Loading, please wait...
Loading, please wait...
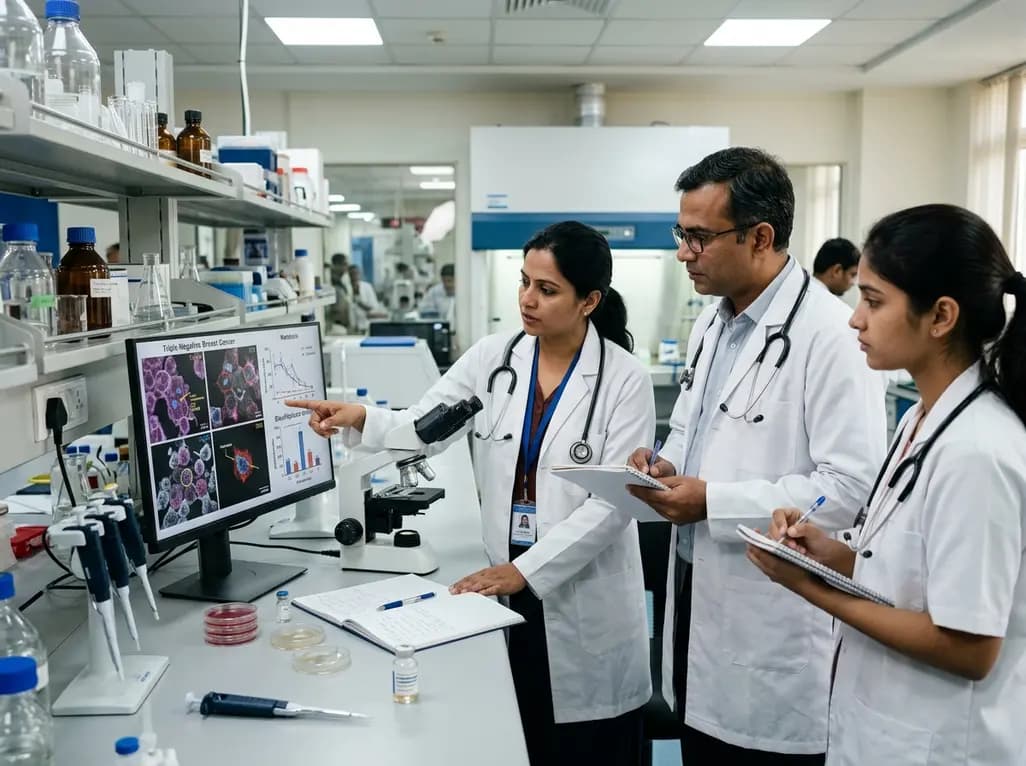
Triple-negative breast cancer remains one of the most challenging malignancies in clinical oncology due to its high heterogeneity and absence of targeted endocrine receptors. Consequently, researchers actively investigate novel molecular dependencies to design effective targeted therapies. Recent scientific discoveries have brought significant attention to the RNA modification machinery, particularly the critical role of YTHDC1 in TNBC. This essential N6-methyladenosine reader dynamically balances nutrient flux under metabolic stress. Thus, unraveling these intricate mechanisms provides valuable clinical insight into overcoming therapeutic resistance and exploiting newly identified cancer vulnerabilities.
Triple-negative breast cancer cells frequently encounter harsh tumor microenvironments characterized by severe nutrient deprivation and elevated oxidative stress. To survive these lethal conditions, malignant cells must rewire their primary metabolic pathways. Emerging molecular research highlights that among all major N6-methyladenosine epigenetic modulators, only the nuclear reader YTHDC1 demonstrates marked, specific overexpression in triple-negative subtypes. Furthermore, elevated expression levels directly correlate with aggressive disease phenotypes and unfavorable patient prognosis.
By functioning as a master epigenetic coordinator, YTHDC1 recognizes methylated transcripts and orchestrates RNA processing to drive adaptive metabolic remodeling. In doing so, this reader protein enables aggressive tumor cells to withstand nutrient scarcity that would otherwise trigger cell death. Moreover, silencing this key regulator markedly impairs tumor viability, which underscores its indispensability in sustaining oncogenic fitness. Consequently, characterizing this nuclear reader sheds light on how RNA methylation directly interfaces with cellular metabolism. Oncology practitioners should recognize that targeting this epigenetic nexus represents an innovative avenue for therapeutic intervention in aggressive breast neoplasms.
Malignant breast tumors exhibit an immense demand for glucose to generate energy and maintain antioxidant defense systems. In triple-negative breast cancer, YTHDC1 directly controls this metabolic demand by stabilizing the messenger RNA of glucose transporter 3, also known as GLUT3. Specifically, YTHDC1 binds to methylated sites on the GLUT3 transcript, which significantly increases its stability and prevents premature degradation.
As a direct result of this stabilization, tumor cells increase glucose uptake from their surroundings. This continuous influx of glucose fuels the pentose phosphate pathway, which generates abundant nicotinamide adenine dinucleotide phosphate, or NADPH. Importantly, intracellular NADPH serves as a critical reducing agent that neutralizes damaging reactive oxygen species. In addition, this continuous supply of reducing equivalents shields cancer cells from oxidative catastrophe during rapid proliferation. Therefore, through post-transcriptional regulation of GLUT3, the nuclear reader maintains continuous bioenergetic flux and protects tumor cells from metabolic exhaustion. Disruption of this axis severely impairs cellular redox capacity, effectively priming aggressive neoplastic cells for selective metabolic destruction.
While glucose uptake provides vital reducing power, excessive cystine accumulation can paradoxically induce a newly characterized, rapid form of cell death termed disulfidptosis. Under normal physiological conditions, activating transcription factor 4, or ATF4, drives the expression of SLC7A11, the cystine-glutamate antiporter. However, YTHDC1 limits the stability of ATF4 mRNA in an m6A-dependent manner within the nucleus. Consequently, this targeted destabilization suppresses the excessive expression of SLC7A11.
By restricting SLC7A11 abundance, the tumor cell carefully controls cystine import and prevents excessive glutamate export. When researchers knock down YTHDC1, ATF4 levels rise rapidly, leading to a marked upregulation of SLC7A11 alongside impaired glucose transport. This imbalance causes intracellular cystine to surge while NADPH levels drop precipitously. Because the cell lacks sufficient NADPH to reduce cystine into cysteine, toxic disulfide bonds accumulate within actin cytoskeleton proteins. Ultimately, this structural collapse triggers lethal disulfidptosis in vitro and in vivo. Thus, YTHDC1 acts as a double-edged regulator that strictly prevents disulfidptosis, preserving cellular integrity in hostile metabolic environments.
Recognizing this unique vulnerability has enabled researchers to develop innovative synthetic biology and nanomedicine strategies. Because nuclear YTHDC1 degrades endogenous ATF4 transcripts before nuclear export, investigators formulated an engineered ATF4 mRNA-targeted nanotherapy. Notably, this lipid nanoparticle system delivers synthetic mRNA directly into the cytoplasm of tumor cells, entirely bypassing nuclear m6A-mediated degradation.
Once translated in the cytoplasm, exogenous ATF4 dramatically upregulates functional SLC7A11 antiporters on the plasma membrane. Consequently, this artificial elevation exposes the tumor cells to severe synthetic lethality when combined with metabolic inhibitors. For instance, pairing this mRNA nanomedicine with the selective GLUT inhibitor BAY-876 induces massive disulfidptosis due to acute NADPH depletion. Similarly, combining ATF4 nanotherapy with the glutaminase inhibitor CB-839 triggers robust cell death driven by severe glutamate deprivation. Furthermore, animal models demonstrate that these combination regimens achieve remarkable tumor regression without noticeable systemic toxicity. Therefore, mRNA-based epigenetic bypass represents a sophisticated paradigm for converting tumor defense mechanisms into therapeutic opportunities.
The discovery of this intricate metabolic network holds profound clinical implications for breast cancer management. Currently, triple-negative breast cancer lacks recurrent actionable surface receptors, which limits clinicians primarily to conventional cytotoxic chemotherapy. However, mapping the YTHDC1-GLUT3-ATF4 axis reveals actionable metabolic dependencies that oncologists can exploit through rational combination therapies.
Moreover, assessing YTHDC1 and SLC7A11 expression patterns may soon serve as predictive biomarkers to identify patient cohorts most likely to respond to metabolic blockade. In clinical settings, integrating targeted mRNA therapeutics with small-molecule metabolic inhibitors offers a dual-pronged strategy to overcome therapeutic resistance. Furthermore, this approach minimizes collateral damage to healthy tissues because non-transformed cells do not exhibit the same extreme metabolic rewiring. In the future, advancing lipid nanoparticle delivery platforms will accelerate the clinical translation of RNA-based metabolic therapies. Ultimately, translating these laboratory discoveries into clinical protocols will empower oncologists to treat refractory triple-negative tumors with greater precision, efficacy, and safety.
YTHDC1 functions as an essential nuclear N6-methyladenosine reader that orchestrates nutrient metabolism and cellular stress responses in triple-negative breast cancer. Specifically, it stabilizes GLUT3 transcripts to enhance glucose uptake and NADPH production while destabilizing ATF4 mRNA to suppress SLC7A11 expression. Consequently, this coordinated regulation prevents metabolic exhaustion, suppresses disulfidptosis, and promotes aggressive tumor progression, making YTHDC1 a key driver of malignancy and an attractive therapeutic target.
SLC7A11 is a transmembrane antiporter that imports extracellular cystine while exporting intracellular glutamate. When SLC7A11 expression increases dramatically under glucose-depleted conditions, cells accumulate high levels of cystine without sufficient NADPH to reduce it to cysteine. Consequently, aberrant disulfide bonds form across actin cytoskeleton filaments, causing severe cytoskeletal collapse and cell death known as disulfidptosis. Thus, manipulating SLC7A11 expression creates a lethal metabolic vulnerability in aggressive cancer cells.
ATF4 mRNA nanotherapy delivers synthetic transcripts directly to the cytoplasm using lipid nanoparticles, effectively bypassing nuclear m6A-mediated degradation orchestrated by YTHDC1. Unlike conventional small molecules that often struggle to selectively modulate intracellular RNA-binding proteins, this mRNA platform restores ATF4 translation directly. Consequently, it forces robust SLC7A11 upregulation and selectively sensitizes tumor cells to metabolic inhibitors like BAY-876 and CB-839, providing higher specificity with reduced systemic toxicity.
Disclaimer: This content is for informational and educational purposes only, and should not be considered medical advice. Always consult a qualified healthcare professional regarding any medical condition or treatment. Refer to the latest local and national guidelines for clinical practice.
References
Lai ZH et al. YTHDC1 Orchestrates Glucose and Glutamate Rewiring to Overcome Lethal Metabolic Stress in Triple-Negative Breast Cancer. Adv Sci (Weinh). 2026 Aug 30. doi: 10.1002/advs.77492. PMID: 42669610.
Liu X, et al. Actin cytoskeleton vulnerability to disulfide stress mediates disulfidptosis. Nat Cell Biol. 2023;25(3):404-414.
Huang S, et al. YTHDC1 promotes triple-negative breast cancer cell proliferation and stemness by regulating BACH1 mRNA stability in an m6A-dependent manner. Mol Cancer Res. 2026;24(8):1158-1172.
Koppula P, et al. Amino acid transporter SLC7A11/xCT at the crossroads of regulating redox homeostasis and nutrient dependency of cancer. Cancer Commun (Lond). 2021;41(1):14-30.
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


YTHDC1 coordinates glucose and glutamate metabolism to protect triple-negative breast cancer cells from metabolic stress and disulfidptosis. Researchers have engineered an ATF4 mRNA nanotherapy that overcomes this axis, unlocking potent combination strategies with metabolic inhibitors.
4 weeks back

A bioengineered oral nanozyme microsupplement converts pathological ROS into a protective polydopamine hydrogel shield. By curbing neutrophil extracellular traps and balancing T cells, this platform outperforms standard 5-ASA in preclinical colitis models, opening new paths for sustained remission.
Today
India experiences an exponential rise in robotic-assisted procedures, with Delhi NCR leading system installations and clinical volume. Discover how multidisciplinary surgical adoption, remote telesurgery breakthroughs, and standardized international consensus guidelines are shaping surgical care nationwide.
Today
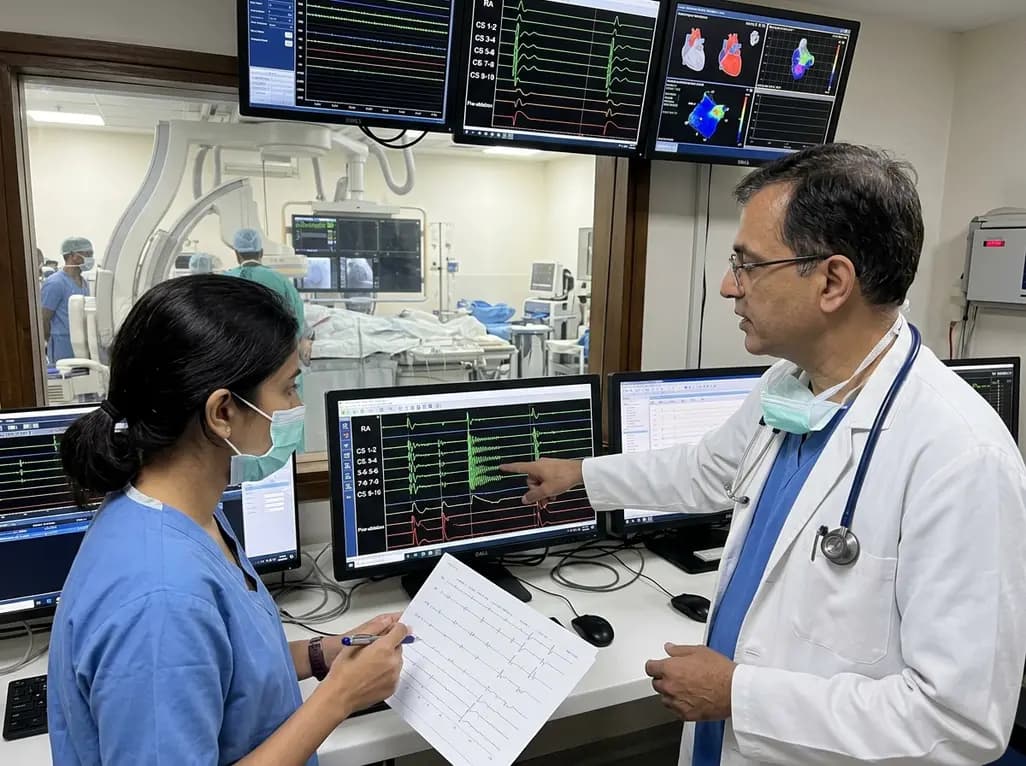
Radiofrequency catheter ablation of the cavotricuspid isthmus successfully terminates typical atrial flutter. This article reviews the mechanisms driving coronary sinus electrogram changes, wavefront activation pathways, and clinical criteria to verify bidirectional block during electrophysiology studies.
Today
A study comparing four knee flexion strength tests after ACL reconstruction using hamstring tendon autografts reveals marked variability in limb symmetry indices. Clinicians must recognize that different strength testing modalities yield divergent return-to-sport clearance rates.
Today
A breakthrough study leverages natural language processing to accurately identify incident medication-related osteonecrosis of the jaw and track antiresorptive drug holidays from clinical narratives in electronic health records, advancing osteoporosis pharmacovigilance and dental safety.
Yesterday