Loading, please wait...
Loading, please wait...
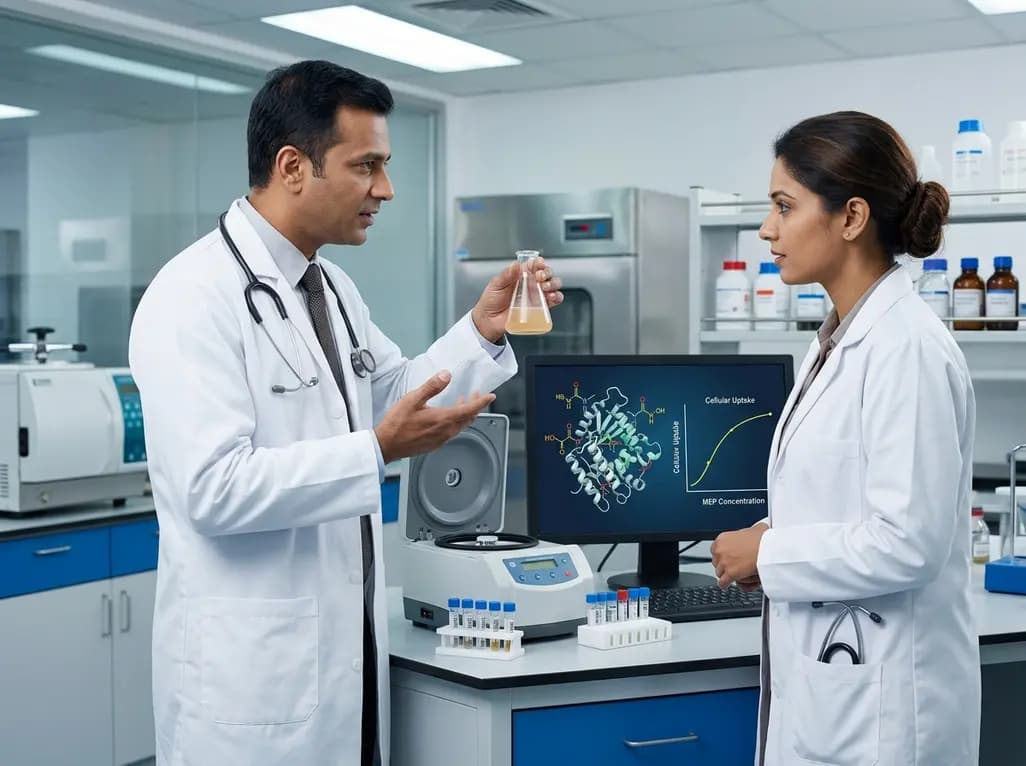
Aspartylglucosaminuria represents an ultra-rare, progressive lysosomal storage disorder that severely impairs neurocognitive and somatic function. The condition arises from pathogenic mutations in the AGA gene, leading to a deficiency in the essential hydrolase aspartylglucosaminidase. Consequently, patients experience an unmitigated intracellular accumulation of glycoasparagines within tissues. Because no disease-modifying pharmacotherapies currently exist, developing a targeted enzyme replacement therapy remains an urgent clinical priority. Recent structural advances offer a promising avenue toward achieving functional therapeutic restoration.
Aspartylglucosaminuria stems from the systemic deficiency of aspartylglucosaminidase, an essential lysosomal amidase. Specifically, this enzyme catalyzes the final step in the catabolism of N-linked glycoproteins. When genetic mutations abolish functional enzyme expression, cytotoxic aspartylglucosamine metabolites accumulate progressively within lysosomes across multiple organ systems. Clinically, affected infants typically present with subtle developmental delays, recurrent respiratory infections, and progressive skeletal alterations. As the storage burden accumulates, patients experience accelerating cognitive decline, loss of speech, motor coordination deficits, and severe neurodegeneration. Furthermore, connective tissue abnormalities lead to coarse facial features, joint hypermobility, and progressive spinal deformities. Because the central nervous system suffers profound damage, early clinical intervention is critical. Unfortunately, traditional supportive care cannot arrest the relentless progression of cellular damage. Thus, restoring active enzyme to affected cells represents the most viable therapeutic strategy. However, delivering active hydrolases across biological membranes into lysosomes presents substantial pharmacological hurdles. Effective therapeutic development requires a deep understanding of natural intracellular trafficking and post-translational maturation processes.
The physiological synthesis and activation of aspartylglucosaminidase involve an intricate series of post-translational modifications. Initially, the AGA gene translates an inactive precursor polypeptide that must fold correctly in the endoplasmic reticulum. Subsequently, this precursor undergoes intramolecular autoproteolytic cleavage to yield separate alpha and beta subunits. These subunits must then assemble into an active heterotetrameric complex to achieve full catalytic competence. In addition, each subunit requires distinct N-linked glycosylation modifications to stabilize the quaternary structure. Without accurate proteolytic cleavage and spatial assembly, the recombinant enzyme fails to attain enzymatic activity. Consequently, previous recombinant manufacturing platforms struggled to produce stable, catalytically mature protein in high yields. Moreover, therapeutic efficacy relies heavily on preserving the structural integrity of both active site pockets. If folding is perturbed, the enzyme rapidly degrades within the extracellular matrix or culture media. Therefore, optimizing expression systems in mammalian host cells is essential to mimic native human maturation. Overcoming these biochemical barriers enables researchers to generate functional, properly folded enzymes suitable for systemic administration.
Targeting therapeutic hydrolases into mammalian lysosomes depends predominantly on the cation-independent mannose-6-phosphate receptor pathway. In native physiology, the Golgi-resident enzyme UDP-N-acetylglucosamine-1-phosphotransferase attaches mannose-6-phosphate residues to newly synthesized lysosomal proteins. However, standard recombinant production systems often produce proteins with suboptimal phosphorylation levels. Consequently, circulating therapeutic enzymes exhibit poor receptor affinity and fail to undergo efficient endocytosis. To resolve this bottleneck, researchers engineered an optimized expression system utilizing a truncated GlcNAc-1-phosphotransferase known as S1S3. This truncated variant retains robust catalytic activity while eliminating regulatory domains that constrain phosphorylation efficiency. By co-expressing S1S3 alongside human aspartylglucosaminidase in HEK293T cells, scientists achieved high-level expression of affinity-tagged, functional recombinant enzymes. Furthermore, the purified enzyme demonstrated high yields directly from conditioned culture media without compromising structural integrity. This novel engineering strategy significantly increases the proportion of phosphorylated oligosaccharide chains on the recombinant protein. As a result, the modified enzyme overcomes historical uptake limitations, facilitating potent cellular entry and targeted lysosomal delivery.
Successful enzyme replacement therapy demands precise glycan engineering to maximize therapeutic bioavailability and intracellular distribution. Experimental investigations confirm that co-expression with the truncated S1S3 phosphotransferase markedly enhances the mannose-6-phosphorylation of aspartylglucosaminidase. Specifically, biochemical characterization revealed that both the alpha and beta subunits require extensive phosphorylation to achieve optimal receptor engagement. When researchers selectively impaired N-glycosylation sites, cellular endocytosis declined dramatically, demonstrating that cellular internalization relies strictly on the mannose-6-phosphate pathway. Moreover, the enhanced phosphorylation pattern directly correlates with accelerated clearance of accumulated substrate in vitro. Because recombinant enzymes with low phosphorylation clear rapidly via hepatic clearance mechanisms, robust mannose-6-phosphate capping prolongs systemic circulation while enhancing targeted endocytosis. Thus, achieving balanced glycosylation and phosphorylation across both subunits is indispensable for maximizing therapeutic potency. In addition, this dual-subunit phosphorylation ensures robust affinity for mannose-6-phosphate receptors across diverse target tissues. Consequently, this engineered biocatalyst demonstrates superior uptake efficiency compared to conventionally manufactured recombinant formulations.
The development of highly phosphorylated recombinant aspartylglucosaminidase brings substantial promise for future clinical management. Although early diagnosis remains challenging due to non-specific initial symptoms, expanded newborn screening programs could identify affected neonates before irreversible neurocognitive decline occurs. Furthermore, early therapeutic administration of optimized enzyme formulations may cross permeable neonatal blood-brain barriers, potentially modifying central neurological outcomes. In older pediatric cohorts, combining systemic enzyme administration with advanced delivery technologies could overcome blood-brain barrier restrictions. In addition, preclinical safety and biodistribution profiles must undergo rigorous validation in transgenic animal models before initiating human clinical trials. Clinicians must also monitor for potential neutralizing antibody formation against the recombinant protein during chronic therapy. Nevertheless, this innovative phosphotransferase engineering platform establishes a scalable blueprint for manufacturing other challenging lysosomal enzymes. Ultimately, translating these preclinical findings into human trials offers renewed hope for patients and families affected by this devastating metabolic disorder.
Aspartylglucosaminuria is an autosomal recessive lysosomal storage disorder caused by pathogenic mutations in the AGA gene. Consequently, deficiency of the enzyme aspartylglucosaminidase impairs the degradation of N-linked glycoproteins. Toxic substrate progressively accumulates in lysosomes throughout the body. Clinically, affected patients experience early developmental delays, recurrent infections, coarse facial features, and progressive cognitive decline. Over time, individuals develop severe intellectual disability, speech impairment, skeletal abnormalities, and progressive motor dysfunction requiring multidisciplinary supportive medical management.
The truncated S1S3 phosphotransferase enhances the attachment of mannose-6-phosphate residues onto newly synthesized aspartylglucosaminidase. Under normal cell culture conditions, recombinant lysosomal enzymes frequently lack sufficient phosphorylation, leading to poor receptor binding. By co-expressing S1S3, researchers significantly augment the phosphorylation of both alpha and beta enzyme subunits. Consequently, the engineered enzyme demonstrates superior affinity for cell-surface mannose-6-phosphate receptors, promoting efficient receptor-mediated endocytosis and accurate intracellular trafficking directly into target lysosomes.
The primary clinical challenge involves delivering adequate concentrations of recombinant enzyme across the blood-brain barrier to reverse neurological pathology. Because aspartylglucosaminuria primarily impacts central nervous system function, systemic enzyme replacement must achieve effective neural biodistribution. Additionally, clinicians must manage potential immune-mediated anti-drug antibody responses resulting from chronic infusions. Early diagnosis through expanded screening is equally critical, as initiating therapy before permanent neurodegenerative damage occurs offers the greatest likelihood of preserving patient cognitive and motor functions.
Disclaimer: This content is for informational and educational purposes only. It is not intended to be a substitute for professional medical advice, diagnosis, or treatment. Always seek the advice of your physician or other qualified health provider with any questions you may have regarding a medical condition. Refer to the latest local and national guidelines for clinical practice.
References
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


Novel glycoengineering advances utilize a truncated phosphotransferase (S1S3) to enhance mannose-6-phosphorylation of aspartylglucosaminidase, significantly boosting cellular uptake and lysosomal delivery for aspartylglucosaminuria therapy.
4 weeks back

Top-down CSF peptidomics uncovers novel endogenous peptide signatures and oxidative modifications differentiating multiple sclerosis from NMOSD, highlighting secretogranin-5 loss as a key biomarker of disease progression.
Today
Fetal foramen ovale restriction typically emerges in the third trimester with right heart enlargement. Evidence demonstrates it does not independently mandate cesarean delivery, and over 93% of postnatal cardiac findings resolve spontaneously within one year, supporting conservative obstetric management.
Today
The latest WHO Global Health Estimates highlight India's life expectancy reaching 71 years alongside a healthy lifespan of 60.6 years. While longevity rebounds post-pandemic, non-communicable conditions now account for three-quarters of top mortalities, underscoring urgent clinical shifts in routine primary care.
Today
New biomechanical research reveals that foot-strike angle significantly influences knee energy absorption during cutting tasks in female soccer players. Understanding these kinematic patterns offers clinicians valuable pathways for targeted ACL injury prevention and athletic retraining.
Today
A comprehensive review analyzes periconceptional GLP-1 receptor agonist exposure, revealing reassuring data on congenital anomalies and adverse perinatal outcomes while emphasizing the need for cautious patient counseling.
Today