Loading, please wait...
Loading, please wait...
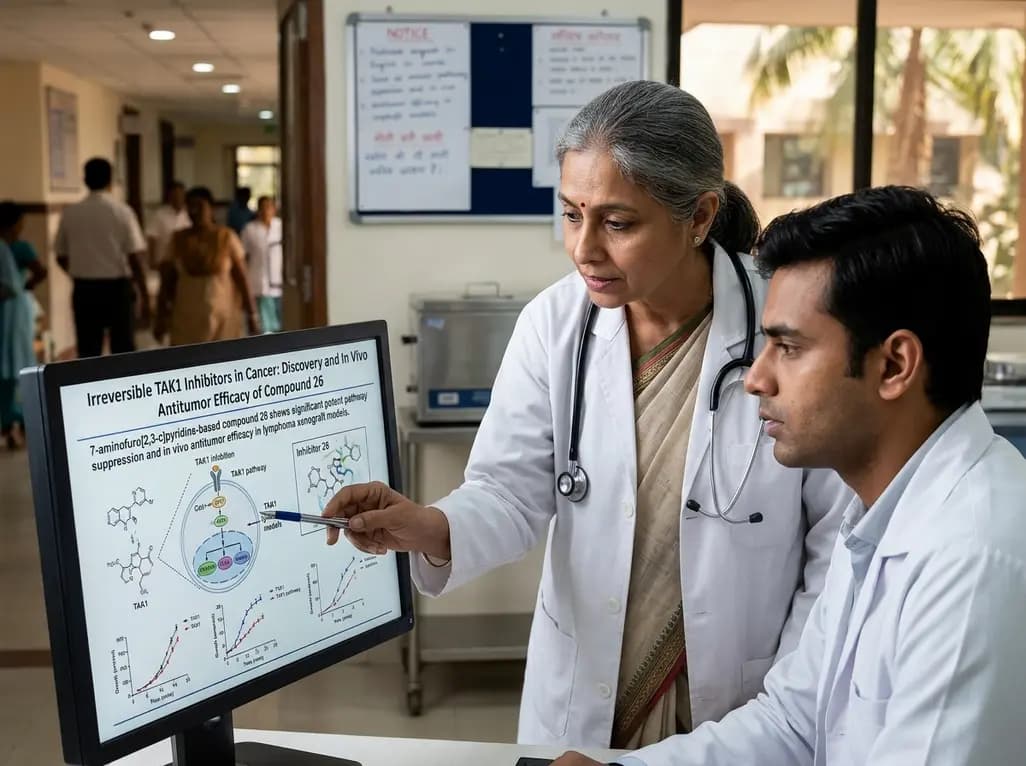
Targeted kinase inhibition remains a cornerstone of precision oncology. Transforming growth factor-β-activated kinase 1 (TAK1) controls vital pro-survival and inflammatory pathways in human malignancies. However, pharmacological targeting of this enzyme has historically faced selectivity hurdles. To overcome these barriers, medicinal chemists recently engineered potent irreversible TAK1 inhibitors. In particular, a novel 7-aminofuro[2,3-c]pyridine derivative offers sustained pathway blockade and notable therapeutic potential.
TAK1 acts as an indispensable signaling junction in modern cellular biology. Specifically, it activates the nuclear factor-kappa B (NF-κB) cascade, which prevents programmed cell death in abnormal cells. In addition, TAK1 directly stimulates mitogen-activated protein kinase pathways, including p38 and JNK cascades. Consequently, these signaling cascades accelerate malignant cell growth and enhance resistance against standard chemotherapeutic agents.
In hematologic cancers, malignant lymphocytes frequently rely on elevated TAK1 signaling to survive metabolic stressors. Furthermore, hyperactive kinase activity protects tumor cells against immune-mediated apoptosis. Therefore, therapeutic disruption of TAK1 can eliminate critical survival advantages in aggressive lymphomas. Historically, small-molecule inhibitors targeting this kinase failed due to low kinome selectivity and rapid in vivo clearance. Moreover, reversible inhibitors often permit rapid pathway reactivation as blood levels fluctuate. Because of these challenges, researchers sought covalent inhibition strategies to secure prolonged enzymatic inactivation. Consequently, oncologists recognize TAK1 as a compelling therapeutic target. Ultimately, targeting TAK1 represents a viable strategy to resensitize resistant cancer cells to apoptotic death.
To address past pharmacologic limitations, medicinal chemists synthesized novel 7-aminofuro[2,3-c]pyridine derivatives. Specifically, they developed compound 26, a specialized molecule bearing an α,β-unsaturated aldehyde warhead. This reactive warhead forms a permanent covalent bond with a conserved cysteine residue near the ATP-binding pocket. Consequently, this covalent interaction locks the kinase in an inactive state and prevents substrate phosphorylation.
Furthermore, the 7-aminofuro[2,3-c]pyridine scaffold provides optimal spatial complementarity within the kinase active site. As a result, compound 26 demonstrates strong nanomolar potency against recombinant TAK1 enzymes. In addition, the covalent mechanism ensures sustained target occupancy even after systemic compound clearance occurs. Importantly, covalent bond formation dramatically increases drug residency time. Notably, earlier covalent kinase inhibitors frequently suffered from unselective protein cross-linking and off-target cytotoxicity. However, compound 26 balances chemical reactivity with precise spatial orientation. Therefore, the compound achieves remarkable target selectivity across diverse cellular assays. Ultimately, this rational design paradigm establishes an encouraging framework for next-generation kinase inhibitors.
In comprehensive cell culture experiments, compound 26 achieved remarkable pathway suppression across multiple tumor models. Specifically, the inhibitor effectively abolished NF-κB transcriptional activity in lymphoma cells. In addition, it suppressed downstream p38 phosphorylation following inflammatory cytokine stimulation. Consequently, compound 26 induced extensive apoptosis, as evidenced by robust caspase activation and cellular fragmentation.
Moreover, the irreversible nature of compound 26 provided sustained pharmacodynamic activity in cell washout experiments. Even after researchers removed the compound from the culture medium, cells maintained durable pathway suppression. Therefore, covalent binding avoids the rapid loss of efficacy that typically hampers reversible kinase inhibitors. Moreover, apoptotic markers increased substantially in a dose-dependent manner. Furthermore, compound 26 preferentially reduced tumor cell viability while sparing non-malignant cell lines. Investigators acknowledged that minor off-target kinase inhibition could contribute slightly to overall cytotoxicity. However, genetic knockdown experiments confirmed that TAK1 inactivation accounts for the observed antitumor effects. Thus, in vitro assays validate compound 26 as a potent and durable molecular disrupter.
Demonstrating consistent in vivo antitumor efficacy remains an essential prerequisite for translational oncology candidates. Notably, researchers evaluated compound 26 in mice bearing human DoHH2 lymphoma xenografts. The covalent inhibitor demonstrated favorable pharmacokinetic distribution and maintained therapeutic tissue exposure throughout the study period. Consequently, systemic administration of compound 26 produced marked, statistically significant tumor growth suppression.
Furthermore, the experimental animals tolerated the therapeutic regimen without significant weight loss or behavioral abnormalities. Tumor histology subsequently revealed widespread apoptotic necrosis alongside reduced cellular proliferation indices. In addition, Western blot analyses of excised tumor tissues confirmed substantial suppression of phosphorylated signaling proteins. Notably, continuous target engagement inhibited tumor vascularization and growth. Therefore, compound 26 successfully penetrated dense tumor tissue and maintained potent pathway inhibition in vivo. These compelling results validate covalent TAK1 targeting as an effective strategy against aggressive lymphoma. Moreover, this study provides vital preclinical proof-of-concept for covalent kinase inhibition in hematologic oncology. Accordingly, compound 26 represents a major scientific advance over previous tool compounds.
The successful preclinical evaluation of compound 26 opens intriguing possibilities for clinical translation. Hematologists frequently encounter aggressive B-cell lymphomas that develop resistance to standard frontline chemotherapy regimens. Because hyperactive survival signaling drives chemoresistance, combining TAK1 inhibitors with conventional cytotoxic drugs could overcome refractory disease states. Furthermore, TAK1 inhibition may synergize with emerging targeted therapies and immune checkpoint antagonists.
However, significant drug development hurdles remain before clinical trial implementation. Specifically, medicinal chemists must optimize the aldehyde warhead to enhance metabolic stability and minimize non-specific protein binding. In addition, ongoing studies must evaluate the potential long-term toxicities associated with sustained systemic TAK1 inhibition. Because TAK1 participates in normal tissue repair and immune surveillance, clinicians must ensure adequate therapeutic windows. Therefore, researchers are designing second-generation analogs with refined pharmacokinetic profiles and improved oral bioavailability. Therefore, cross-disciplinary collaboration will drive these translational efforts forward. Ultimately, compound 26 serves as an indispensable tool compound that accelerates the rational design of clinical-grade targeted anticancer therapeutics.
TAK1 serves as a critical signaling hub that coordinates pro-inflammatory pathways, cell survival, and resistance to apoptosis. In several hematologic and solid malignancies, abnormal TAK1 activation drives persistent NF-κB and MAPK signaling, allowing cancer cells to proliferate and resist conventional chemotherapy. Therefore, selectively inhibiting TAK1 disrupts these essential survival cascades, sensitizes malignant cells to death ligands like TNF-α, and induces apoptosis, offering a compelling strategy for targeted cancer therapies.
Traditional reversible kinase inhibitors bind non-covalently to target enzymes and dissociate rapidly when plasma drug concentrations decline. In contrast, covalent irreversible inhibitors form a permanent chemical bond with specific amino acid residues, such as cysteine, within the catalytic pocket. Consequently, this irreversible binding provides sustained target suppression, prolongs pharmacodynamic activity, and reduces the required dosing frequency. Furthermore, this covalent mechanism prevents rapid therapeutic wash-out, maintaining prolonged pathway blockade even within hostile tumor microenvironments.
Although compound 26 demonstrates promising in vivo antitumor activity, several developmental hurdles remain before human trials begin. Specifically, medicinal chemists must refine the reactive aldehyde warhead to eliminate off-target proteomic reactivity and reduce systemic toxicity risks. Additionally, researchers must optimize pharmacokinetic properties, including oral bioavailability, systemic clearance rates, and plasma protein binding. Finally, rigorous safety toxicology studies must evaluate long-term impacts on immune homeostasis and normal tissue maintenance across diverse animal models.
Disclaimer: This content is for informational and educational purposes only... Refer to the latest local and national guidelines for clinical practice.
References
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


Researchers have discovered compound 26, a 7-aminofuro[2,3-c]pyridine-based irreversible inhibitor of TAK1 that demonstrates potent pathway suppression and significant in vivo antitumor efficacy in lymphoma xenograft models, presenting a promising avenue for targeted antineoplastic therapy.
Today
Researchers demonstrate that central induction of synthetic torpor in non-hibernating rats provides striking cardioprotection, reducing myocardial infarct size by ~40% during ischaemia-reperfusion injury through endogenous survival signaling, independent of hypothermia.
Today
Ascertainment bias in cancer case-control studies inflates odds ratios and penetrance estimates when cases are drawn from high-risk clinics. A landmark modeling study demonstrates substantial bias ranging up to 23.72, highlighting the critical need for ascertainment adjustment in genetic risk counseling.
Today
Researchers have engineered novel light-on photoswitchable adrenoceptor agonists targeting β1-receptors. By remaining inactive until illuminated, these precision molecules offer reversible optical control, tailored relaxation kinetics, and distinct efficacy profiles, paving the way for targeted cardiovascular therapies.
Today
Chronic nonbacterial osteomyelitis (CNO) is a pediatric autoinflammatory bone disorder. This review covers the French protocol, emphasizing whole-body MRI as the diagnostic gold standard, prudent biopsy avoidance, first-line NSAID treatment, and second-line biologic or bisphosphonate therapies for spinal lesions.
Today
A novel fluorine-functionalized near-infrared copper fluorescent probe selectively illuminates lung tissue, revealing critical links between hyperglycemia, ER stress, and pulmonary malignancy for improved early diagnosis.
Today