Loading, please wait...
Loading, please wait...

Highly effective CFTR modulator therapy has dramatically reshaped the clinical management and long-term prognosis of cystic fibrosis. The introduction of elexacaftor/tezacaftor/ivacaftor, known as ETI, restored approximately forty percent of physiological CFTR chloride transport in individuals with at least one F508del mutation. Consequently, thousands of patients experienced marked improvements in lung function, nutritional status, and overall quality of life. However, clinicians increasingly recognize that genetic heterogeneity complicates therapeutic outcomes. Specifically, individuals who harbor the cis-acting complex allele L467F;F508del fail to achieve meaningful clinical improvements after starting ETI. This rare complex variant combines the classical F508del deletion with a secondary missense mutation in the first nucleotide-binding domain. As a result, the double-mutant protein undergoes severe conformational instability and biosynthetic arrest. Because standard triple therapy cannot overcome this structural defect, these patients face persistent disease progression and therapeutic exclusion. Therefore, discovering novel pharmacotherapies represents an urgent clinical priority for respiratory specialists and pediatricians worldwide.
To overcome this treatment barrier, investigators evaluated the next-generation triple regimen comprising vanzacaftor, tezacaftor, and deutivacaftor, abbreviated as VTD. Vanzacaftor functions as a structurally distinct, next-generation CFTR corrector designed to stabilize mutant proteins through alternative molecular binding pockets. Furthermore, deutivacaftor provides enhanced pharmacokinetic stability over first-generation potentiators, allowing once-daily dosing regimens. In this groundbreaking study, researchers compared the biological efficacy of ETI against VTD using highly differentiated primary human nasal epithelial cultures. These cultures originated from seven individuals with cystic fibrosis carrying at least one L467F;F508del complex allele alongside nine healthy control subjects. By measuring transepithelial short-circuit currents in Ussing chambers, the investigators quantified CFTR-mediated chloride secretion under highly physiological ex vivo conditions. Importantly, patient-derived nasal epithelia accurately mirror bronchial epithelial transport properties. Thus, this robust translational model provides invaluable predictive data regarding how novel small molecules rescue defective channel function across diverse patient genotypes.
The experimental electrophysiological findings clearly demonstrate significant functional differences between the two triple-drug regimens. Treatment of patient-derived nasal epithelia with ETI yielded only a modest, non-significant numerical shift in chloride currents. In stark contrast, treatment with VTD stimulated statistically significant chloride secretion following cyclic AMP activation with forskolin and IBMX. Moreover, addition of the specific CFTR inhibitor CFTRinh-172 confirmed that restored transepithelial currents directly resulted from functional CFTR channels. Although VTD provided only partial rescue relative to wild-type healthy controls, the absolute functional increase represents a critical biological threshold. Clinical studies consistently establish that even modest restorations of channel transport, often between ten and twenty percent of normal function, produce meaningful clinical gains in pulmonary function and sweat chloride reduction. Consequently, this study confirms that vanzacaftor possesses unique pharmacodynamic properties capable of overcoming structural conformational blocks that completely derail earlier correctors like elexacaftor.
These findings highlight the expanding role of personalized theratyping in complex genetic pulmonary diseases. Because complex alleles remain rare within global cystic fibrosis populations, randomized clinical trials cannot easily recruit sufficient cohorts for conventional phase three protocols. Therefore, ex vivo evaluation using primary respiratory epithelial cells serves as an essential surrogate tool to guide individualized clinical decisions. When clinicians encounter apparent treatment failure or unexpectedly poor sweat chloride reduction under standard regimens, comprehensive genetic sequencing often reveals undetected cis-variants. Furthermore, patient-derived epithelia reliably predict whether alternative pharmacological combinations can salvage channel gating and trafficking. By validating the efficacy of VTD in this difficult-to-treat genotype, the research establishes a reproducible pathway for compassionate access and off-label therapeutic trials. In addition, these results reassure clinicians that non-responsiveness does not always reflect irreversible end-stage tissue damage, but rather an allele-specific pharmacological mismatch.
The validation of VTD against the L467F;F508del complex allele offers practical lessons for pulmonologists, general practitioners, and pediatric providers managing respiratory tract disorders. First, clinicians must thoroughly investigate unexpected non-responders who possess the common F508del mutation. Identifying secondary cis-mutations like L467F through full-gene sequencing prevents inappropriate long-term exposure to ineffective regimens. Second, as regulatory agencies approve once-daily vanzacaftor-based triple combinations, physicians gain a potent therapeutic alternative for refractory complex genotypes. Nevertheless, practitioners must remember that restoring chloride secretion ex vivo represents only the initial step toward comprehensive disease management. Patients transitioning to novel modulators still require rigorous pulmonary toilet, monitoring of hepatic transaminases, and nutritional optimization. Ultimately, the successful rescue of previously refractory complex alleles underscores a promising shift toward precision medicine, ensuring that every affected individual can access life-changing genetic correction.
The L467F;F508del complex allele occurs when two distinct genetic variations arise on the same CFTR gene strand in cis configuration. In addition to the common F508del phenylalanine deletion, a leucine-to-phenylalanine substitution occurs at codon 467. Consequently, this secondary structural alteration causes profound misfolding and severe biosynthetic processing arrest. Standard therapies fail to overcome this conformational defect, rendering affected patients non-responsive to first-generation triple combinations.
VTD succeeds because vanzacaftor features a novel chemical scaffold that binds alternative regions on the mutant CFTR channel. Unlike elexacaftor, which cannot correct the severe dual misfolding caused by both mutations, vanzacaftor stabilizes the protein during early ribosomal synthesis. Therefore, vanzacaftor facilitates membrane trafficking and membrane insertion. When paired with tezacaftor and deutivacaftor, it partially restores channel gating and apical chloride secretion ex vivo.
Clinicians should suspect complex alleles whenever patients carrying documented F508del mutations display no clinical or sweat chloride improvements after starting standard ETI therapy. In such cases, physicians must order comprehensive CFTR gene sequencing rather than basic panel testing to identify cis-variants. Furthermore, specialized centers can perform ex vivo nasal epithelial testing to evaluate whether switching to a vanzacaftor-based regimen effectively restores functional ion transport.
Disclaimer: This content is for informational and educational purposes only... Refer to the latest local and national guidelines for clinical practice.
References
Berger J et al. Vanzacaftor/tezacaftor/deutivacaftor partially restores CFTR function of the L467F;F508del complex allele in primary nasal epithelial cells. J Cyst Fibros. 2026 Oct 03. doi: undefined. PMID: 42829305.
Pedemonte N, Tomati V, Capurro V, et al. Vanzacaftor-Tezacaftor as an alternative therapeutic resource for the ETI-Resistant L467F-F508del Allele: Ex vivo prediction and exploratory clinical assessment. J Cyst Fibros. 2026; doi: 10.1016/j.jcf.2026.05.015. PMID: 42243036.
Sondo E, Cresta F, Pastorino C, et al. The L467F-F508del Complex Allele Hampers Pharmacological Rescue of Mutant CFTR by Elexacaftor/Tezacaftor/Ivacaftor in Cystic Fibrosis Patients. J Pers Med. 2022;12(5):789. doi: 10.3390/jpm12050789.
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


Novel ex vivo evidence demonstrates that the next-generation triple combination vanzacaftor/tezacaftor/deutivacaftor partially rescues CFTR channel function in patients harboring the treatment-refractory L467F;F508del complex allele, overcoming non-responsiveness to elexacaftor/tezacaftor/ivacaftor.
Today
A pharmacometric and machine learning study reveals that specific vaginal bacteria alter cervical antiretroviral exposure, highlighting the need for precision HIV PrEP strategies.
Yesterday
Abdominal aortic aneurysm rupture remains a catastrophic vascular emergency. Groundbreaking research reveals that USP53 accelerates aneurysm progression by reprogramming smooth muscle cell metabolism toward aerobic glycolysis, highlighting a promising target beyond conventional diameter-based surveillance.
Today
A breakthrough study leverages natural language processing to accurately identify incident medication-related osteonecrosis of the jaw and track antiresorptive drug holidays from clinical narratives in electronic health records, advancing osteoporosis pharmacovigilance and dental safety.
Today
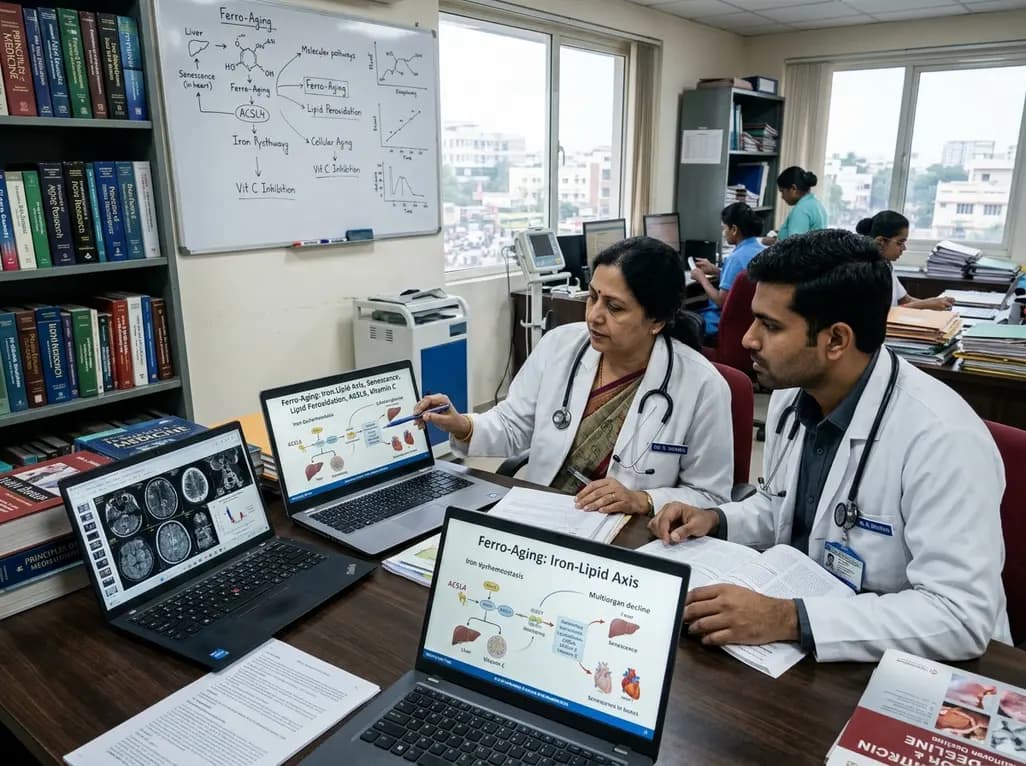
Recent research defines ferro-aging as a conserved iron-lipid axis driving organ decline across primates. Age-related iron dyshomeostasis stimulates ACSL4-mediated lipid peroxidation, promoting cellular senescence. Notably, vitamin C directly inhibits ACSL4, suppressing phospholipid oxidation and multi-organ decay.
Yesterday