Loading, please wait...
Loading, please wait...
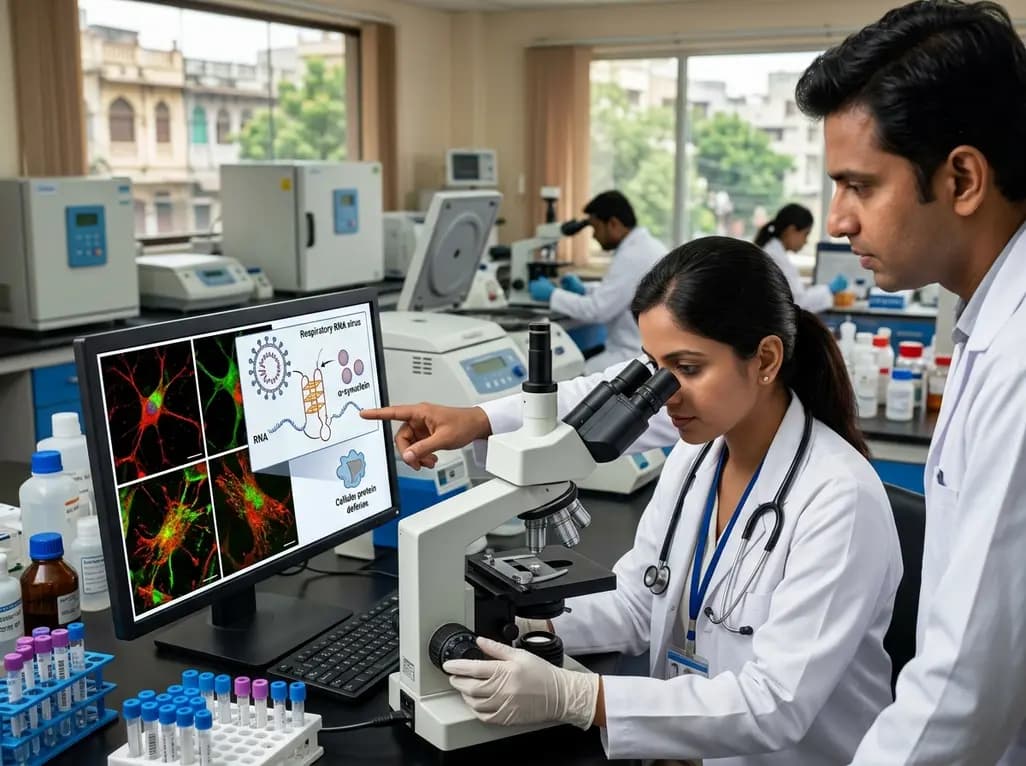
Understanding the biological triggers of neurodegeneration remains one of modern medicine's greatest challenges. Clinicians have long noted neurological sequelae following acute viral infections. Now, compelling experimental evidence illuminates the Parkinson's disease viral link. Researchers from the Centre for Cellular and Molecular Biology in Hyderabad explored this critical relationship. Specifically, their findings show how transient viral infections trigger long-term neuropathology. The team demonstrated that viral genetic material directly accelerates toxic protein clustering. Furthermore, these biomolecular interactions promote rapid alpha-synuclein amyloidogenesis in host neurons. Consequently, these discoveries bridge the longstanding gap between acute systemic infections and chronic neurodegenerative decline.
Neurodegenerative disorders such as Parkinson's disease feature toxic amyloid aggregates within the central nervous system. Under normal physiological conditions, alpha-synuclein regulates synaptic vesicle trafficking and neurotransmitter release. However, environmental stressors, cellular insults, or somatic mutations can destabilize this native protein. When destabilization occurs, alpha-synuclein misfolds and forms soluble oligomeric species. Eventually, these oligomers assemble into insoluble fibrillar structures known as Lewy bodies.
These toxic aggregates disrupt axonal transport, impair mitochondrial respiration, and induce synaptic failure. Moreover, misfolded alpha-synuclein spreads across interconnected neural circuits via prion-like seeding. Consequently, neurons experience unrelenting oxidative stress and neuroinflammation. Ultimately, dopaminergic neurons in the substantia nigra undergo apoptosis, driving hallmark motor symptoms. While genetic mutations explain select familial cases, idiopathic disease dominates clinical practice. Therefore, environmental factors, including neurotropic and systemic viruses, warrant detailed investigation. Cohort studies have repeatedly highlighted post-viral parkinsonian symptoms following major epidemics. Until now, however, clinicians lacked precise molecular frameworks explaining how viral agents drive this proteinaceous cascade. In particular, physician-scientists have sought tangible biomolecular evidence directly linking viral components to protein aggregation. Furthermore, identifying the initial biophysical triggers of alpha-synuclein clumping remains pivotal for targeted intervention.
To decipher this pathological connection, CCMB scientists examined viral genetic architecture inside host cells. Respiratory pathogens, including influenza A and SARS-CoV-2, utilize RNA genomes that fold into complex shapes. Specifically, these viral genomes fold into secondary structures termed RNA G-quadruplexes. These distinct configurations comprise planar guanine quartets held together by robust Hoogsteen hydrogen bonds. During active viral replication, the cytoplasm rapidly accumulates high concentrations of these structural motifs.
Importantly, the researchers discovered that viral RNA G-quadruplexes bind directly to human alpha-synuclein. This binding event triggers liquid-liquid phase separation and drives rapid sol-gel transition of the protein. As a result, alpha-synuclein molecules condense swiftly into insoluble amyloid fibrils. Viral RNA G-quadruplexes effectively act as catalytic seeds, dramatically shortening normal aggregation lag times. Furthermore, this nucleating event does not require permanent viral persistence within neurons. Instead, even transient exposure to viral genetic material leaves lasting structural alterations. Thus, an acute respiratory infection can initiate autonomous amyloidogenesis that persists long after systemic recovery. Accordingly, this finding offers a biological foundation for post-infectious neurodegeneration. Indeed, this catalytic mechanism illustrates how foreign nucleic acids compromise intrinsic proteostatic stability. Consequently, even mild respiratory viral encounters might accelerate early subclinical protein misfolding events.
Fortunately, host cells mobilize intrinsic enzymatic safeguards against invading pathogens. In this context, the study spotlighted DDX39A, an essential DEAD-box RNA helicase. Under resting conditions, DDX39A resides inside the cell nucleus to coordinate mRNA processing. However, upon viral infection, DDX39A rapidly redistributes into the cytoplasm. Once in the cytosolic space, DDX39A targets both viral RNA G-quadruplexes and monomeric alpha-synuclein.
Through ATP hydrolysis, DDX39A unwinds the folded viral RNA G-quadruplexes with remarkable efficiency. Consequently, the dismantled viral genome cannot support efficient replication, causing intracellular viral titers to drop. Simultaneously, this helicase activity removes the RNA scaffold driving alpha-synuclein phase condensation. Thus, DDX39A acts as a dual-function guardian that restricts infection while preventing amyloid clumping.
Nevertheless, this cellular defense entails a delicate biomolecular trade-off. Interestingly, cooperative binding with alpha-synuclein enhances the helicase's unwinding capacity. Yet, when viral RNA burdens overwhelm host resources, this protective equilibrium falters. As a result, the cellular DDX39A pool becomes saturated and depleted. Therefore, massive viral loads leave alpha-synuclein unprotected, accelerating irreversible amyloid deposition. Ultimately, differences in individual helicase reserves may determine neurological vulnerability. Furthermore, these insights clarify why clinical outcomes differ markedly among infected individuals.
A vital insight from this discovery involves the relationship between infection frequency and neurodegenerative risk. Typically, a single viral bout does not directly trigger clinical Parkinson's disease. Healthy cells deploy robust quality control networks, such as molecular chaperones and the autophagy-lysosome system. Consequently, healthy neurons effectively dissolve small oligomeric clusters and restore physiological proteostasis.
However, repeated viral infections may gradually overwhelm these endogenous repair systems. Frequent bouts of influenza, COVID-19, or related RNA viruses cause recurring surges of viral G-quadruplexes. Each successive encounter draws DDX39A out of the nucleus and stresses cytoplasmic defenses. Furthermore, advancing chronological age naturally degrades basal proteostatic capacity and reduces helicase reserves. Therefore, repeated viral challenges can produce cumulative biomolecular damage over several decades.
Similarly, recurrent infections stimulate chronic microglial activation and systemic neuroinflammation. Over time, persistent inflammatory mediators disrupt the blood-brain barrier and exacerbate synaptic injury. As a result, each infection acts as an incremental secondary hit within vulnerable brain regions. In genetically susceptible individuals, these recurrent molecular insults lower the clinical threshold for neurodegeneration. Ultimately, lifelong viral burden may explain why certain patients develop idiopathic synucleinopathies. Accordingly, evaluating an individual's cumulative infectious history could offer valuable diagnostic clues for clinicians.
These molecular findings carry profound clinical implications for modern medical practice. Primarily, they underscore the neuroprotective value of rigorous viral disease prevention. For instance, broad vaccination coverage against influenza and SARS-CoV-2 reduces severe infections and systemic viral dissemination. Moreover, early antiviral administration during acute illnesses limits the accumulation of viral RNA G-quadruplexes. By dampening viral replication early, clinicians may prevent the initiation of pathological alpha-synuclein nucleation.
In addition, this research highlights novel molecular avenues for therapeutic development. Pharmaceutical scientists can design small molecules that stimulate or stabilize DDX39A helicase activity. Such agents would reinforce host cellular defenses against viral replication and protein aggregation simultaneously. Alternatively, synthetic compounds could block binding sites between viral RNA and alpha-synuclein. Consequently, disrupting this initial phase transition represents a transformative neuroprotective strategy.
Furthermore, physicians should recognize post-viral neurological manifestations during routine outpatient follow-up. While post-infectious screening protocols remain under development, tracking cognitive and motor signs remains prudent. Understanding the molecular link between viral exposure and amyloid formation will undoubtedly reshape preventative neurology. Ultimately, targeted interventions targeting RNA-protein interactions may preserve long-term cognitive and motor health. Indeed, bridging virology and neurodegenerative therapeutics creates unprecedented opportunities for clinical medicine.
Q1: How do viral RNA structures accelerate alpha-synuclein clumping in Parkinson's disease?
RNA viruses fold their genomic material into stable three-dimensional configurations termed RNA G-quadruplexes. When these viral structures accumulate in the cytoplasm during acute infections, they bind directly to alpha-synuclein. This binding event triggers rapid liquid-liquid phase separation and drives the sol-gel transition of alpha-synuclein. Consequently, the viral RNA behaves as a catalytic scaffold that accelerates the formation of insoluble amyloid fibrils, which form classic Lewy bodies in Parkinson's disease.
Q2: What protective function does the DDX39A protein perform during viral infection?
DDX39A is an essential DEAD-box RNA helicase normally located in the cell nucleus. Upon viral invasion, DDX39A redistributes into the cytoplasm and binds both viral RNA G-quadruplexes and alpha-synuclein. Using ATP hydrolysis, the enzyme unwinds these folded viral structures, which significantly suppresses viral replication and lowers intracellular viral load. Additionally, this enzymatic unwinding dismantles the RNA scaffold, thereby actively slowing down the aberrant phase transition and amyloid formation of alpha-synuclein.
Q3: Does every respiratory viral infection lead to the development of Parkinson's disease?
No, an individual viral infection does not inevitably trigger Parkinson's disease. Cellular proteostatic mechanisms and helicase enzymes successfully resolve transient amyloidogenesis and eliminate viral particles in most instances. However, repeated respiratory infections across a lifetime can deplete protective cellular reserves and overburden clearance pathways. Consequently, recurrent exposures combined with genetic vulnerability and advancing age can incrementally increase an individual's lifetime risk for developing progressive neurodegenerative synucleinopathies.
Disclaimer: This content is for informational and educational purposes only. It does not constitute medical advice or replace professional judgment. Refer to the latest local and national guidelines for clinical practice.
References
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


A pioneering study from CSIR-CCMB Hyderabad demonstrates how respiratory RNA viruses like influenza and SARS-CoV-2 promote alpha-synuclein amyloid aggregation through RNA G-quadruplex structures, while identifying nuclear helicase DDX39A as a vital cellular defense mechanism.
Today
Data from the 2026 Workforce Health Index reveals that 38.7% of urban employees and dependents exhibit abnormal glycemic control. Emerging at increasingly younger ages, this metabolic shift intersects with severe vitamin deficiencies, highlighting the urgent need for early workplace screenings and targeted clinical care.
Today
The Maharashtra Food and Drug Administration has enforced stringent compliance orders under food safety laws. The directive prohibits commercial promotions, infant formula samples, gifts, and clinician sponsorships while mandating explicit breastfeeding notices and rapid adverse event reporting across healthcare facilities.
Today
Haryana Chief Minister Nayab Singh Saini has rolled out landmark performance-linked financial incentives for public sector clinicians and allied staff. The comprehensive reform introduces free intravitreal diabetic retinopathy management, flexible empanelment for private specialists, and digital infrastructure upgrades.
Today
A cross-sectional study reveals a strong positive correlation between problematic internet use and kinesiophobia in adults aged 18-45 with non-specific low back pain, highlighting how digital overuse reinforces movement fear and complicates musculoskeletal recovery.
Today
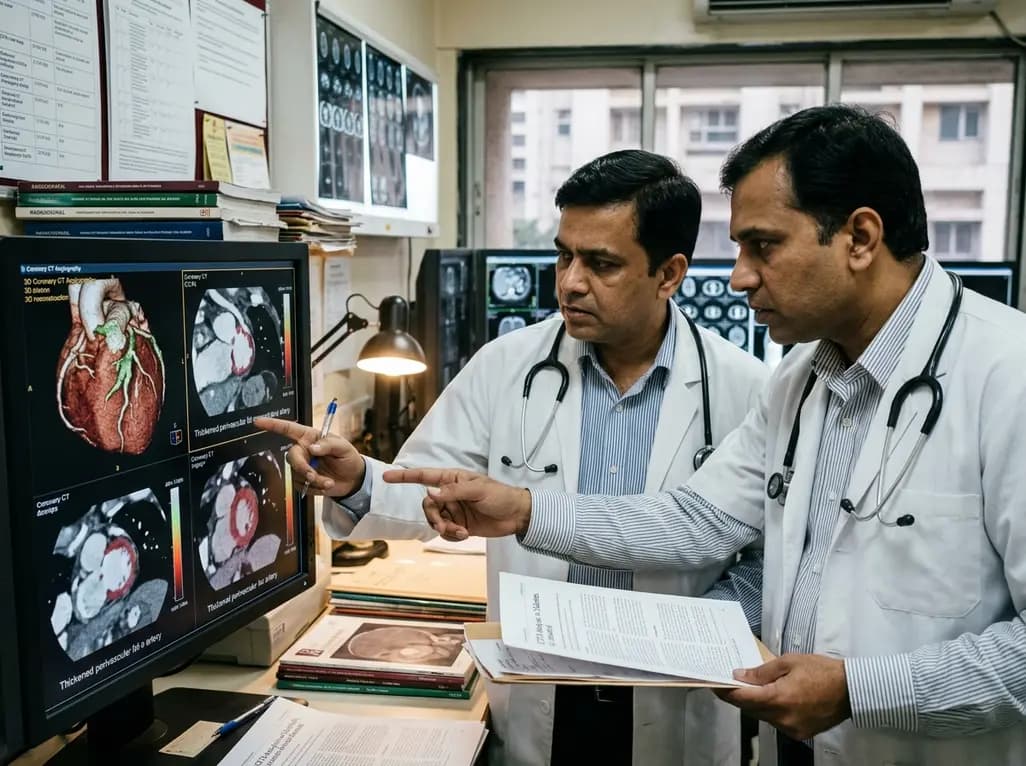
A recent coronary CT angiography study demonstrates that diabetic patients exhibit accelerated plaque burden, impaired CT-FFR, and elevated perivascular fat attenuation index values, reflecting heightened vascular inflammation that worsens across adverse metabolic phenotypes.
Today