
Loading, please wait...
Loading, please wait...
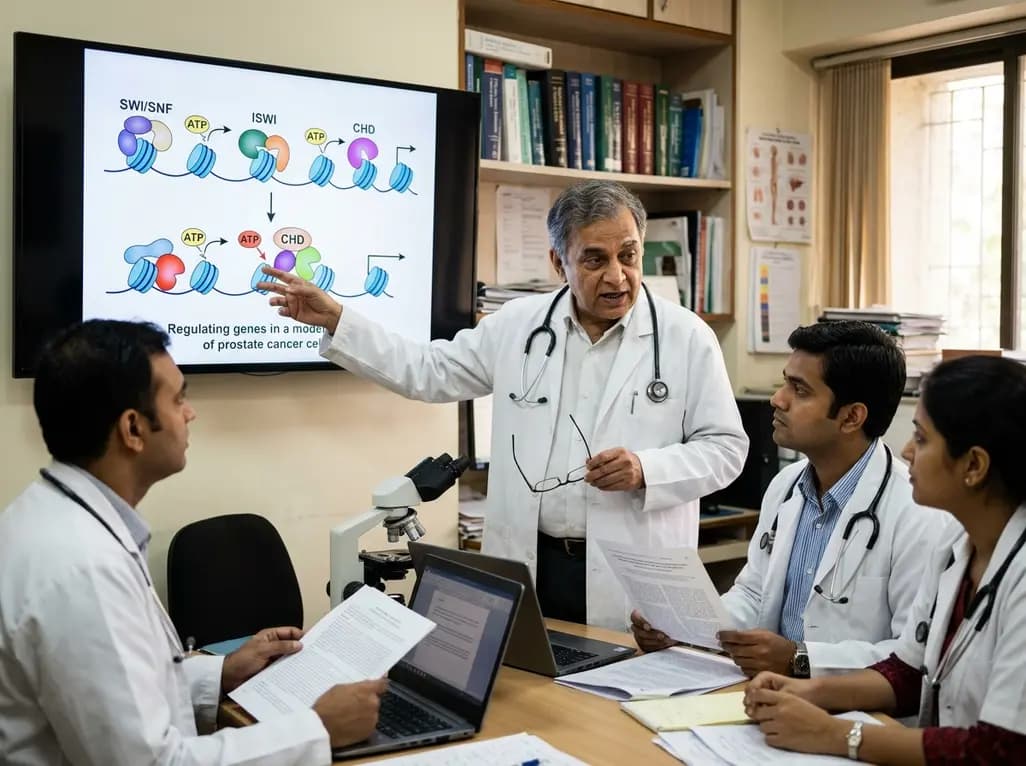
Prostate malignancy remains one of the leading contributors to male cancer mortality across the globe. Although androgen receptor signaling inhibitors offer substantial survival gains, acquired resistance frequently emerges in advanced stages. Emerging evidence indicates that epigenetic plasticity governs these adaptive cellular transitions. In particular, ATP-dependent chromatin remodelers in prostate cancer orchestrate nucleosome organization across the entire genome. These macromolecular complexes modulate chromatin accessibility, dictating where oncogenic transcription factors bind. Consequently, understanding chromatin remodelers provides pivotal insights into treatment failure and illuminates innovative therapeutic avenues for aggressive disease.
Mammalian cells organize ATP-dependent chromatin remodelers into four specialized families. These core families comprise SWI/SNF, ISWI, CHD, and INO80/SWR complexes. Each family utilizes catalytic ATPase subunits that hydrolyze ATP to reposition, eject, or restructure nucleosomes. For instance, SWI/SNF complexes slide and eject octamers to establish nucleosome-depleted regions at distal enhancers. In contrast, ISWI and CHD family members assemble regularly spaced nucleosomal arrays. Furthermore, INO80 and SWR complexes specialize in histone variant exchange, specifically modulating H2A.Z dynamics during transcription and DNA repair.
In malignant prostate tissue, these families coordinate the epigenomic landscape that controls transcription factor access. As prostate adenocarcinoma progresses, altered subunit expression shifts chromatin architecture. Therefore, cooperative activity between distinct remodelers governs cellular lineage identity. When tumor cells experience therapeutic pressure, aberrant remodeler recruitment permits continuous transcription of survival genes. Thus, structural diversity explains why chromatin remodelers in prostate cancer produce multifaceted phenotypic consequences.
The androgen receptor functions as the premier transcriptional driver throughout prostate ontogenesis. However, the receptor cannot access condensed chromatin without dynamic epigenetic assistance. SWI/SNF complexes, particularly canonical BAF assemblies, facilitate receptor recruitment to regulatory DNA elements. These remodelers loosen tightly packed chromatin at lineage-specific enhancers, allowing coregulators like FOXA1 and HOXB13 to bind cooperatively.
Nevertheless, continuous androgen receptor inhibition drives extensive epigenomic adaptation. Resistant tumors frequently amplify androgen receptor expression or express truncated splice variants like AR-V7. In addition, mutated chromatin complexes sustain enhancer accessibility under androgen-depleted conditions. For example, loss of CHD1 reorganizes receptor binding profiles across oncogenic enhancers, promoting enzalutamide resistance. Similarly, SWI/SNF ATPase subunits maintain super-enhancer looping architecture that preserves androgen-regulated gene expression. Consequently, chromatin remodeling complexes act as indispensable gatekeepers that preserve transcriptional signaling despite potent antiandrogen therapies.
Advanced prostate tumors harbor recurrent genomic alterations in classical tumor suppressor genes. Specifically, combined loss of PTEN, TP53, and RB1 triggers rapid disease acceleration. Interestingly, these genetic lesions do not simply disable cell cycle checkpoints. Instead, they actively install novel chromatin remodeling dependencies that clinicians can therapeutically exploit.
For instance, PTEN deficiency hyperactivates downstream oncogenic signaling while establishing a distinct vulnerability to SWI/SNF inhibition. Cells lacking functional PTEN rely heavily on intact SWI/SNF ATPase catalytic activity to manage transcriptional stress and DNA replication. When researchers inhibit these ATPase enzymes in PTEN-null models, malignant cells undergo selective apoptosis. Similarly, simultaneous inactivation of TP53 and RB1 alters chromatin compaction, creating collateral sensitivity to chromatin-directed interventions. Consequently, synthetic lethality provides an exceptional framework for precision oncology strategies. By targeting induced dependencies created by tumor suppressor deficiency, oncologists can eradicate treatment-refractory cells while sparing normal tissues.
Under intense pharmacological suppression, prostate carcinomas increasingly escape through lineage plasticity. Tumors transition away from luminal differentiation toward neuroendocrine prostate cancer or double-negative phenotypes devoid of androgen and neuroendocrine markers. Intriguingly, ATP-dependent chromatin remodelers govern this profound epigenetic reprogramming.
During lineage shifts, complex composition undergoes dramatic structural remodeling. For example, canonical BAF complexes exchange accessory subunits, altering genomic localization toward embryonic loci. Furthermore, SMARCA4 interacts with alternative master transcription factors such as TCF7L2, fueling pro-proliferative WNT pathway activity in double-negative subtypes. This selective rewiring enables malignant cells to survive complete androgen receptor downregulation. In addition, aberrant chromatin remodeling silences luminal differentiation genes while simultaneously opening promoters for neuronal transcriptional networks like ASCL1. As a result, chromatin remodeling complexes dictate cellular identity, directly sustaining drug resistance through lineage transformation.
Given their central role in treatment resistance, chromatin remodeling complexes represent highly attractive therapeutic targets. Historically, designing small-molecule inhibitors against multimeric chromatin assemblies presented severe structural challenges. However, the advent of proteolysis-targeting chimeras, or PROTAC degraders, has revolutionized this landscape.
Novel dual SMARCA2 and SMARCA4 degraders, such as AU-15330, show remarkable efficacy in preclinical models. These degraders compact chromatin rapidly at vital enhancers, dislodging androgen receptors, MYC, and FOXA1 from DNA. Moreover, these molecules achieve striking tumor regression in enzalutamide-resistant castration-resistant prostate cancer without intolerable toxicity. Parallel drug discovery programs actively pursue small-molecule inhibitors directed against ISWI and INO80 complexes to exploit homologous recombination deficiencies. In the future, clinical oncologists will likely combine these chromatin-directed degraders with existing antiandrogens or immunotherapies. Therefore, targeting chromatin remodelers in prostate cancer establishes a transformative therapeutic paradigm for overcoming fatal therapy resistance.
Chromatin remodelers alter nucleosome spacing across regulatory enhancer regions, ensuring continuous genomic access for the androgen receptor. Even during potent antiandrogen therapy, aberrant remodeler activity sustains transcriptional activation of critical oncogenes. Furthermore, these complexes facilitate the binding of truncated androgen receptor variants and alternative pioneer factors. Consequently, tumors maintain downstream survival pathways, rendering standard hormonal therapies ineffective and driving progression to castration-resistant disease.
Loss of the tumor suppressor PTEN hyperactivates oncogenic pathways, resulting in elevated transcriptional replication stress within malignant cells. To survive this heightened genomic tension, PTEN-deficient cells depend heavily on SWI/SNF ATPase activity to remodel chromatin and maintain gene expression. Therefore, pharmacological inhibition or degradation of SWI/SNF catalytic subunits selectively triggers catastrophic chromatin collapse in PTEN-null cells. As a result, malignant prostate cells undergo apoptosis while normal cells remain largely unaffected.
Traditional small-molecule inhibitors often fail to abolish remodeler function completely because multi-subunit chromatin complexes possess extensive non-enzymatic scaffolding properties. In contrast, targeted PROTAC degraders induce rapid proteasomal destruction of essential catalytic subunits like SMARCA2 and SMARCA4. Consequently, PROTACs dislodge entire oncogenic transcriptional complexes from chromatin, disassembling super-enhancers that sustain tumor growth. This targeted degradation achieves profound anti-tumor responses in treatment-resistant prostate cancer while circumventing common enzymatic resistance mechanisms.
Disclaimer: This content is for informational and educational purposes only and does not constitute medical advice, diagnosis, or treatment recommendations. Clinical decisions should always be made by qualified healthcare professionals based on individual patient assessment and prevailing clinical guidelines. Refer to the latest local and national guidelines for clinical practice.
References
Torres LM et al. ATP-Dependent Chromatin Remodelers in Prostate Cancer Progression and Therapeutic Resistance. Endocrinology. 2026 Sep 16. doi: undefined. PMID: 42747431.
Xiao L et al. Targeting SWI/SNF ATPases in enhancer-addicted prostate cancer. Nature. 2022;601(7893):434-439.
Thienger P et al. A Double-Negative Prostate Cancer Subtype Is Vulnerable to SWI/SNF-Targeting Degrader Molecules. Cancer Discov. 2026;16(4):e1-e18.
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.
ATP-dependent chromatin remodelers (SWI/SNF, ISWI, CHD, and INO80) regulate nucleosome dynamics to govern androgen receptor signaling and lineage plasticity in prostate cancer. Targeting these complexes, especially in PTEN-deficient and double-negative subtypes, opens vital therapeutic frontiers.
last month
Psoriasis is a chronic systemic inflammatory condition requiring comprehensive clinical intervention beyond topical care. Explore evidence-based insights into systemic comorbidities, patient quality of life, and targeted biologic options such as IL-23 inhibitors for sustained disease control.
Today
A nationwide cohort study published in JAMA Pediatrics reveals that maternal exposure to wildfire fine particulate matter, especially in late pregnancy, is linked to an elevated risk of childhood autism. Clinicians should evaluate ambient air quality risks and provide proactive protective counseling to mothers.
Today
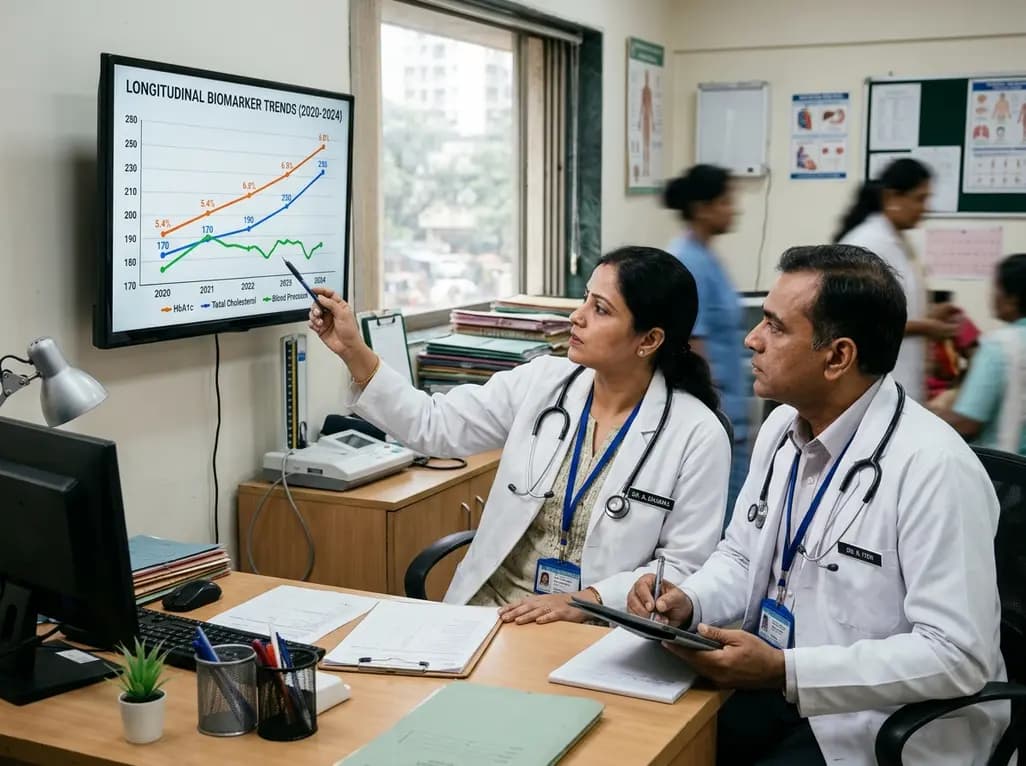
Annual health checks only offer static snapshots. Real-world evidence from 2.1 million insured Indians reveals that tracking dynamic biomarker trends over time enables clinicians to spot evolving cardiometabolic risks early, reverse borderline pathology, and deliver tailored preventive interventions.
Today
Modern 3D anthropometry reveals that pediatric leg growth ceases years before spinal elongation, exposing the clinical illusion of the classical Anderson-Green model. Learn how segmental growth dissociation improves surgical timing for epiphysiodesis in adolescent leg length discrepancy.
2 days back
With packaged goods giant HUL backing FSSAI's proposed front-of-pack red warning labels for foods high in fat, sugar, and salt, India takes a major step toward curbing non-communicable diseases. Learn how clearer nutrition disclosures empower clinicians, inform consumer choices, and drive dietary reform.
Today