Loading, please wait...
Loading, please wait...
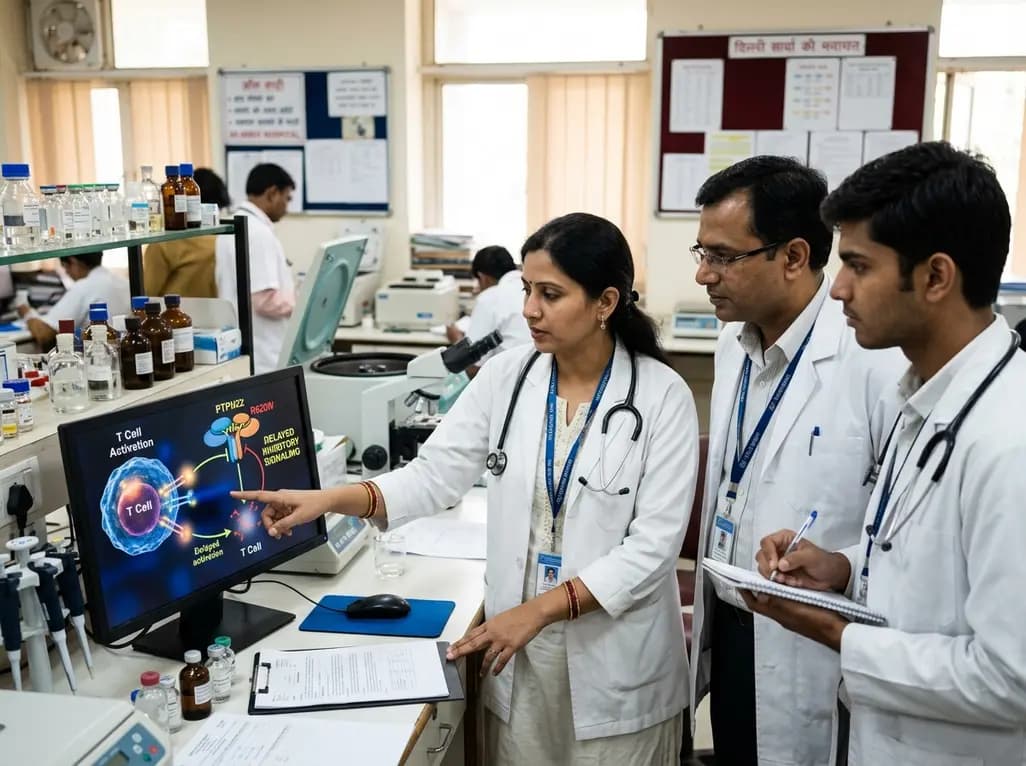
Precise control of adaptive immunity requires balanced positive triggers and timely repressive signals. In this regulatory framework, PTPN22 T cell activation pathways maintain self-tolerance and prevent unintended tissue damage. Protein tyrosine phosphatase non-receptor type 22, commonly known as PTPN22 or LYP, serves as a crucial lymphoid-specific gatekeeper. When naive T cells engage cognate antigens presented by major histocompatibility complexes, phosphorylation cascades initiate downstream effector functions. However, unrestrained activation generates chronic systemic inflammation and triggers tissue destruction. PTPN22 actively dephosphorylates key early activation kinases, including Lck, Fyn, and ZAP70. Consequently, this enzymatic action dampens early signaling cascades before excessive clonal expansion occurs. Classical paradigms previously described PTPN22 solely as an immediate early dampener. Recent experimental data revise this dogma by highlighting a coordinated, delayed inhibitory cascade. By coordinating negative feedback loops, the phosphatase ensures that antigen clearance occurs without triggering persistent autoimmunity. Therefore, understanding this phosphatase offers indispensable insights into how human immune cells calibrate responses against self-antigens.
Groundbreaking molecular analyses have identified an intricate multimolecular architecture that governs signal attenuation. Rather than acting as an isolated monomer, PTPN22 forms a specialized inhibitory complex with critical cytoplasmic partners. Specifically, mass spectrometry and biochemical imaging confirm interactions between PTPN22, C-terminal Src kinase, STS-1, and PSTPIP. During the earliest phases of antigen recognition, T cells must maintain robust signaling to decide cell fate. Therefore, cells intentionally postpone the recruitment of this inhibitory machinery. As activation progresses over several minutes, these four regulatory proteins coalesce into dense supramolecular structures. Within this protective cluster, Csk phosphorylates inhibitory residues while PTPN22 removes activating phosphate moieties from key signaling intermediates. Furthermore, PSTPIP acts as an adaptor scaffolding element that anchors the entire assembly to the cytoskeleton. STS-1 reinforces this inhibition by blocking ubiquitin-mediated survival cascades. Together, these cooperative partners build an enzymatic barrier that shuts down hyperactive intracellular communication. Consequently, the delayed assembly allows initial immune triggering while guaranteeing timely signal cessation.
Advanced high-resolution microscopy demonstrates that spatial organization dictates the precise timing of immune suppression. When a T cell encounters an antigen-presenting cell, surface receptors form submicron domains known as microclusters. These microscopic assemblies coordinate essential early signaling events and cytokine secretion. Initially, active tyrosine kinases concentrate within these microclusters without encountering significant phosphatase opposition. However, live-cell imaging reveals that PTPN22 and its regulatory partners migrate toward these microclusters with distinct delayed kinetics. This temporal pause ensures that necessary activation signals cross the required threshold before negative feedback commences. Once the inhibitory complex arrives, it disrupts microcluster stability and drives receptor internalization. Moreover, the delayed migration prevents premature signal abort during productive infections. By timing this recruitment precisely, the cell creates a self-limiting signaling window. Knockout mouse models devoid of these clustering molecules display persistent, uninhibited microcluster phosphorylation. Consequently, these knockout cells fail to terminate signaling, leading to pathological hyperreactivity and prolonged inflammatory responses.
Genetic studies have long associated the single-nucleotide polymorphism rs2476601 with multiple autoimmune disorders. This functional variant causes an arginine-to-tryptophan substitution at position 620 within the non-catalytic domain of PTPN22. Structural analyses demonstrate that this alteration disrupts the crucial proline-rich motif responsible for protein-protein interactions. Specifically, the R620W mutant exhibits dramatically reduced binding affinity for Csk and STS-1. Because of this defective interaction, the inhibitory complex cannot assemble correctly at the immunological synapse. As a direct consequence, the delayed negative feedback loop completely collapses in carrying individuals. T cells expressing the mutant variant fail to silence antigen receptor microclusters in a timely manner. Instead, these lymphocytes sustain prolonged phosphorylation cascades and generate excessive inflammatory cytokines, including interleukin-2 and interferon-gamma. Furthermore, impaired complex formation weakens central and peripheral tolerance checkpoints. Autoreactive clones escape thymic deletion and initiate destructive reactions against host tissues. Thus, this single amino acid change transforms a vital inhibitory rheostat into a permissive driver of human autoimmunity.
For practicing physicians managing autoimmune conditions, these molecular findings clarify long-standing clinical paradoxes. The PTPN22 R620W polymorphism represents one of the strongest non-HLA genetic risk factors for rheumatoid arthritis, type 1 diabetes, and systemic lupus erythematosus. Patients harboring this risk allele frequently present with aggressive disease trajectories and refractory inflammation. Furthermore, understanding delayed feedback kinetics explains why baseline immune markers may appear normal despite severe episodic flares. When these patients encounter environmental triggers or infections, their T cells overreact and produce exaggerated cytokine bursts. Clinicians should recognize that genetic susceptibility directly alters signaling duration rather than simple on-off switching. In addition, routine genotyping may eventually help stratify patients who are prone to hyperactive T cell responses. Recognizing individual molecular risk profiles empowers clinicians to personalize therapeutic monitoring and anticipate organ complications. Therefore, incorporating pathway dynamics into clinical assessments offers a sophisticated perspective on disease heterogeneity across diverse patient cohorts.
Uncovering the delayed negative feedback machinery opens promising avenues for therapeutic innovation in rheumatology and clinical immunology. Traditional immunosuppressive agents broadly suppress lymphocyte proliferation, frequently causing serious opportunistic infections. In contrast, modern drug discovery focuses on restoring endogenous regulatory complexes to achieve selective immunomodulation. Pharmacological modulators that stabilize interactions between PTPN22, Csk, and STS-1 could rescue defective negative feedback in genetic carriers. Alternatively, small molecules targeting downstream kinases can shorten microcluster lifespan in hyperactive T cells. In oncology, transiently blocking PTPN22 assembly might conversely boost anti-tumor immune responses by sustaining T cell activation against resistant malignancies. Preclinical mouse models already show that modulating this phosphatase axis reshapes immune responses without complete systemic immunosuppression. Consequently, translating these structural discoveries into clinical practice could revolutionize treatment algorithms for autoimmune disorders. Ultimately, restoring natural inhibitory timing promises precise disease control while preserving vital defensive immunity against pathogenic threats.
PTPN22 regulates T cell activation by assembling an inhibitory multimeric complex with Csk, STS-1, and PSTPIP minutes after antigen stimulation. Initially, T cells require robust receptor phosphorylation to initiate immune defenses. Subsequently, this regulatory complex migrates to T cell receptor microclusters, where it dephosphorylates critical activating kinases. Consequently, this delayed spatial recruitment prevents premature signal termination while ensuring that immune responses shut down appropriately before causing systemic tissue damage.
The R620W missense mutation alters a key proline-rich binding motif in PTPN22, significantly reducing its physical interaction with Csk and STS-1. As a direct result, the delayed inhibitory complex fails to assemble at receptor microclusters following T cell activation. Without timely feedback inhibition, autoreactive T cells exhibit prolonged kinase activation and exaggerated inflammatory cytokine production, driving chronic tissue destruction in rheumatoid arthritis, lupus, and type 1 diabetes.
This discovery provides clinicians with a precise mechanistic framework explaining why PTPN22 variant carriers experience heightened disease severity and therapeutic resistance. Instead of relying exclusively on non-specific immunosuppressive drugs, future clinical therapies may specifically stabilize the inhibitory complex or downregulate hyperactive microcluster signaling. Furthermore, genetic screening for the R620W allele can help physicians identify patients who require earlier intervention and tailored biologics to prevent irreversible inflammatory joint and organ damage.
Disclaimer: This content is for informational and educational purposes only... Refer to the latest local and national guidelines for clinical practice.
References
Hashimoto-Tane A et al. PTPN22 mediates delayed negative feedback of T cell activation by forming an inhibitory signaling complex. Int Immunol. 2026 Sep 25. doi: undefined. PMID: 42786963.
Begovich AB, et al. A missense single-nucleotide polymorphism in a gene encoding a protein tyrosine phosphatase (PTPN22) is associated with rheumatoid arthritis. Science. 2004;305(5681):256-258.
Bottini N, et al. A functional variant of lymphoid tyrosine phosphatase is associated with type 1 diabetes. Nat Genet. 2004;36(4):337-339.
Stanford SM, Bottini N. PTPN22: the archetypal autoimmune gene. Nat Rev Rheumatol. 2014;10(10):602-610.
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


New immunological research reveals how PTPN22 forms a delayed inhibitory signaling complex to restrain T cell activation, explaining how the autoimmune-linked R620W variant disrupts feedback control and promotes persistent inflammation in conditions like rheumatoid arthritis and type 1 diabetes.
Today
A landmark cluster randomized trial demonstrates that community-based mass dog vaccination achieves superior coverage and sustains protective herd immunity against rabies far better than conventional campaign models, offering vital lessons for global and Indian elimination targets.
Today
Explore the crucial link between dietary protein and immunocompetence. This review examines how low-protein nutrition shifts metabolic resources away from pathogen defense toward early reproduction, compromising host survival and immunity against severe bacterial infections.
Today
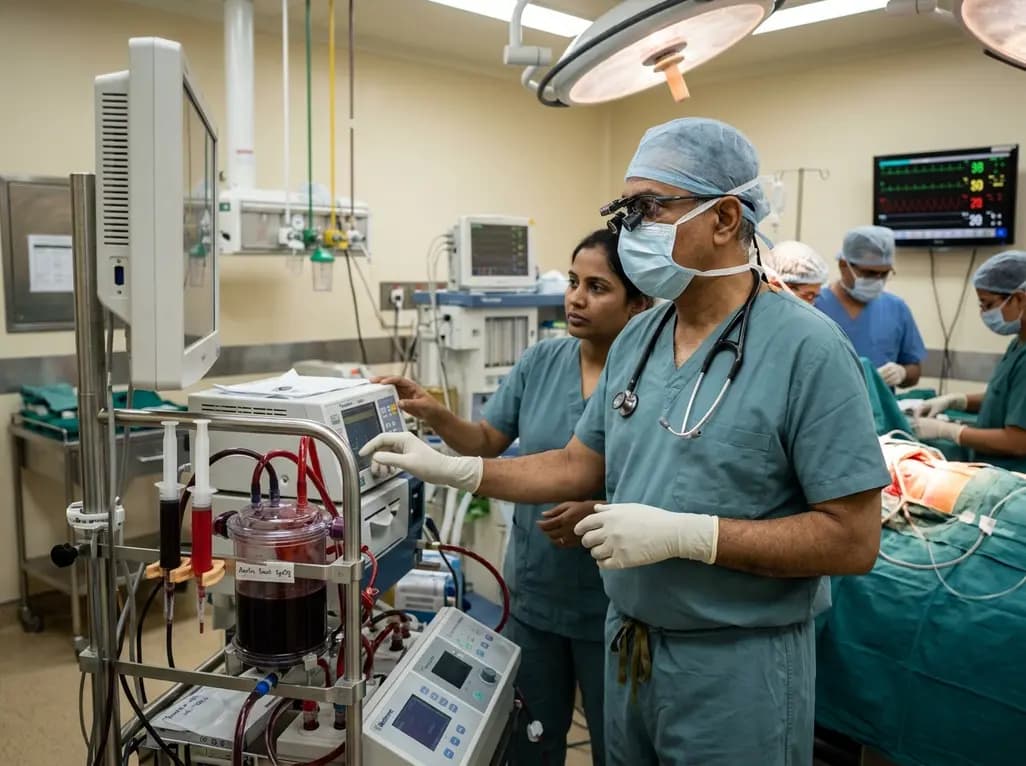
Differential oxygenation during peripheral TA-NRP can cause undetected coronary hypoxemia due to arch vessel clamping. A novel circuit with continuous aortic root monitoring detects desaturation early, facilitating left ventricular unloading and preventing allograft dysfunction and unnecessary heart discard.
Today
A longitudinal study reveals that patients undergoing staged bilateral hip arthroscopy experience progressive contralateral labral deterioration during surgical intervals. However, structural degradation does not compromise 2-year postoperative clinical outcomes or patient-reported satisfaction scores.
Today
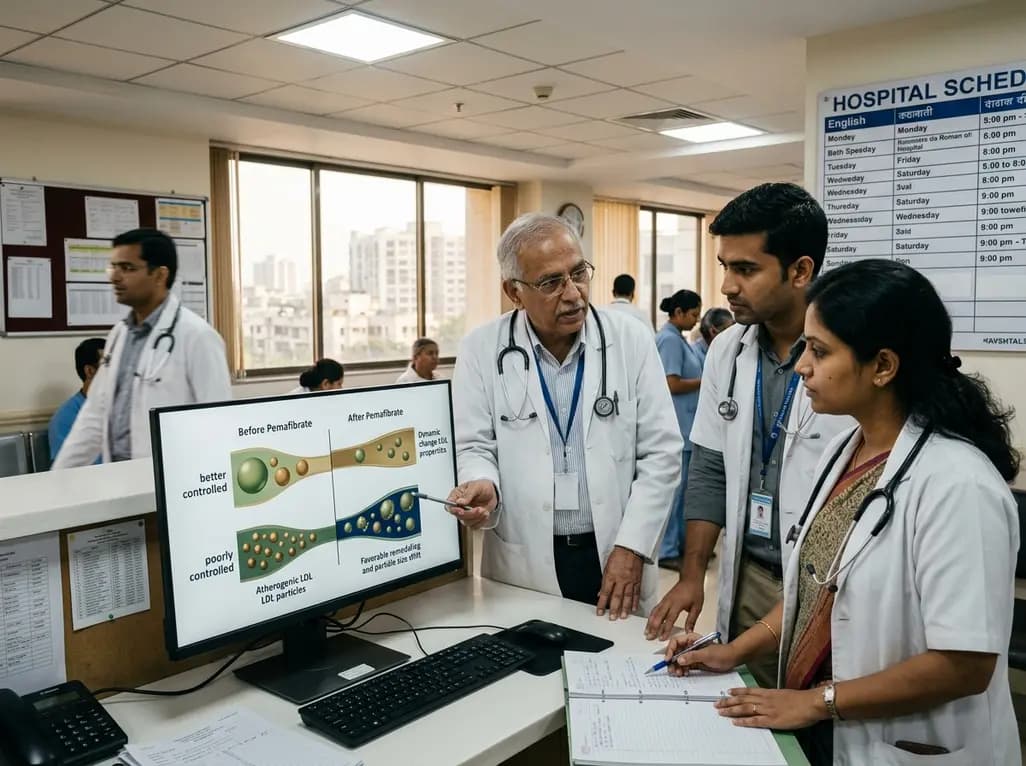
A post hoc analysis of the PARM-T2D study shows pemafibrate exerts baseline-dependent effects on LDL-C and particle size in type 2 diabetes and hypertriglyceridemia. It lowers triglycerides and remnants while favorably remodeling atherogenic LDL particles based on initial lipid control status.
Today