Loading, please wait...
Loading, please wait...
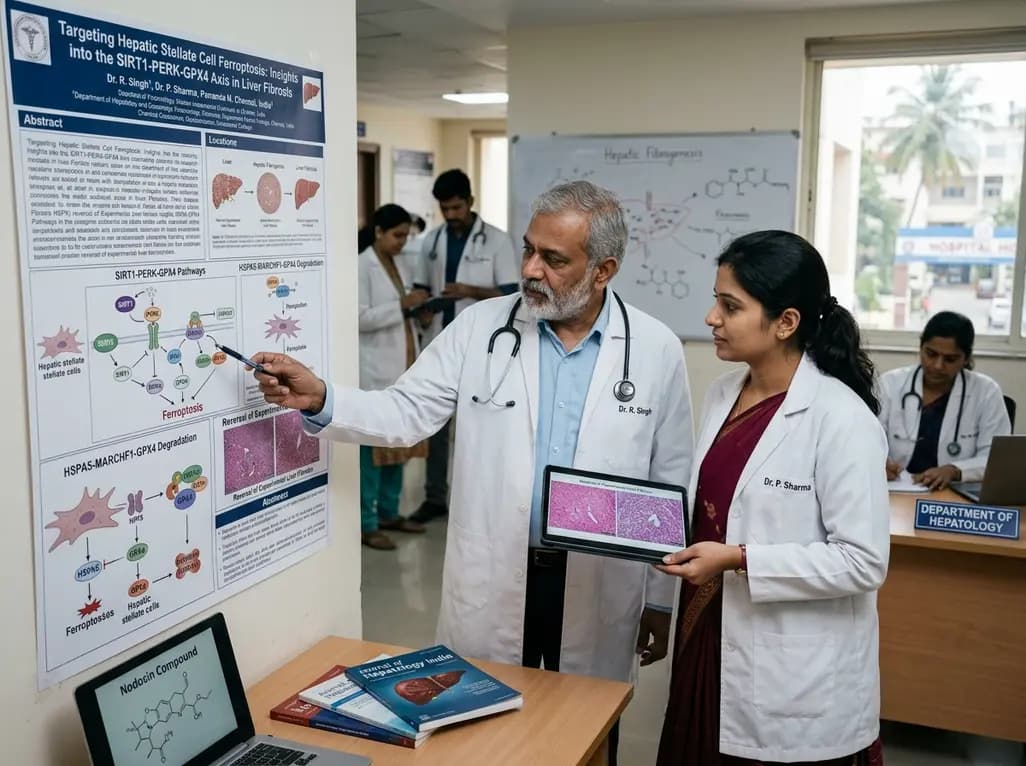
Liver fibrosis represents a major clinical challenge across global gastroenterology and hepatology practices. Chronic hepatic injury consistently triggers the transdifferentiation of quiescent perisinusoidal cells into collagen-secreting myofibroblasts. Consequently, inducing hepatic stellate cell ferroptosis has emerged as a promising strategy to halt or reverse fibrotic matrix accumulation. A groundbreaking study published in Antioxidants & Redox Signaling identifies an endoplasmic reticulum stress-proteostasis axis that governs this cellular vulnerability.
Persistent hepatic inflammation activates quiescent hepatic stellate cells into active myofibroblasts. Consequently, these transformed cells synthesize excessive extracellular matrix components, including collagen types I and III. Although traditional therapies attempt to suppress activation signals, selectively eliminating activated myofibroblasts offers a more definitive therapeutic endpoint. Ferroptosis represents an iron-dependent form of non-apoptotic cell death characterized by lethal lipid peroxide accumulation. Furthermore, glutathione peroxidase 4 serves as the central guardian preventing iron-mediated lipid peroxidation. When cellular GPX4 activity declines, toxic lipid reactive oxygen species overwhelm endogenous defenses. As a result, the activated stellate cell undergoes rapid plasma membrane rupture and cell death. Preclinical evidence confirms that clearing these matrix-producing cells directly attenuates hepatic architectural distortion. In contrast, quiescent hepatocytes largely remain protected from these selective ferroptotic triggers. Therefore, delineating the exact proteostatic machinery controlling GPX4 stability provides actionable insights for rational anti-fibrotic drug development.
Endoplasmic reticulum stress serves as a pivotal regulator of cellular adaptation during chronic liver injury. When unfolded proteins accumulate within the lumen, the sensory kinase PERK undergoes activation and autophosphorylation. Subsequently, phosphorylated PERK phosphorylates eukaryotic initiation factor 2 alpha, selectively enhancing activating transcription factor 4 translation. Researchers demonstrated that nodosin, an ent-kaurane diterpenoid from Isodon serra, actively disrupts this pathway. Mechanistically, nodosin enhances sirtuin 1 activity, which deacetylates PERK at lysine residue 1101. Because deacetylation suppresses aberrant PERK activation, downstream ATF4 expression drops significantly in treated stellate cells. Consequently, this transcriptional downregulation diminishes protective chaperone production within the endoplasmic reticulum. This molecular finding proves that post-translational deacetylation directly modulates kinase activity under stress conditions. Moreover, suppressing ATF4 alters downstream transcriptional programs essential for cell survival. Clinicians should note that modulating this checkpoint selectively sensitizes myofibroblasts to metabolic exhaustion without disrupting baseline hepatocyte viability.
Heat shock protein family A member 5, also recognized as BiP or GRP78, acts as an essential chaperone. Under basal conditions, ATF4 drives HSPA5 expression to maintain proteostatic equilibrium. However, when nodosin suppresses the PERK-ATF4 axis, HSPA5 levels fall precipitously. Importantly, the authors demonstrated that HSPA5 physically shields GPX4 from recognition by membrane-associated ring-CH-type finger 1. MARCHF1 functions as an E3 ubiquitin ligase that targets specific lysine residues on client proteins. Once HSPA5 dissociates from the protective complex, MARCHF1 gains direct access to GPX4. Specifically, MARCHF1 mediates K48-linked polyubiquitination of GPX4 at the critical lysine 191 residue. As a direct consequence, the 26S proteasome rapidly degrades ubiquitinated GPX4. Without functional GPX4 to neutralize toxic lipid hydroperoxides, intracellular lipid peroxides surge uncontrollably. Thus, the HSPA5-MARCHF1 molecular toggle establishes GPX4 proteostasis as the primary gatekeeper controlling ferroptotic sensitivity in fibrogenic cells.
To evaluate in vivo relevance, the investigators established carbon tetrachloride-induced hepatic fibrosis in C57BL/6J mice. Following two weeks of repeated toxic injury, animals received oral nodosin at graduated doses of 1, 5, or 10 mg/kg daily. Histopathological analysis revealed marked reductions in bridging fibrosis, pseudolobule formation, and total collagen deposition. Furthermore, serum transaminase levels decreased substantially in nodosin-treated cohorts, indicating preserved parenchymal function. Quantitative assays confirmed elevated markers of lipid peroxidation exclusively within the stromal compartment of treated livers. Concurrently, immunohistochemistry demonstrated reduced alpha-smooth muscle actin and diminished GPX4 expression in non-parenchymal zones. These observations confirm that pharmacological intervention effectively promotes target cell clearance in established disease. Additionally, nodosin demonstrated excellent oral bioavailability and systemic tolerability across all experimental cohorts. Therefore, natural diterpenoids targeting specific proteostatic nodes present viable therapeutic leads for advanced translational testing in chronic liver injury.
Chronic liver disease represents an escalating healthcare burden across India and worldwide. While lifestyle modification and antiviral regimens address primary etiologies, direct anti-fibrotic pharmacotherapies remain exceptionally limited. Targeting hepatic myofibroblast clearance via ferroptosis provides an innovative paradigm that complements existing etiology-directed strategies. Moreover, identifying the SIRT1-PERK-HSPA5-MARCHF1 axis uncovers multiple distinct druggable targets within a single regulatory circuit. Clinicians can envision future small-molecule therapeutics designed to enhance MARCHF1 ligase activity or selectively inhibit HSPA5-GPX4 interaction. Furthermore, combination regimens incorporating natural diterpenoids alongside standard metabolic interventions may achieve superior histological regression. Nevertheless, rigorous clinical trials must confirm target selectivity to prevent off-target lipid peroxidative damage in healthy hepatic parenchymal tissue. As translational research progresses, mapping endoplasmic reticulum proteostasis will undoubtedly refine personalized risk stratification and anti-fibrotic drug screening.
Glutathione peroxidase 4 functions as an indispensable phospholipid hydroperoxidase that reduces toxic lipid hydroperoxides into non-toxic lipid alcohols. In activated hepatic stellate cells, sustained GPX4 expression prevents lethal lipid reactive oxygen species accumulation. Consequently, destabilizing GPX4 compromises cellular redox defense, which triggers rapid iron-dependent cell death and attenuates pathological extracellular matrix secretion.
Nodosin enhances SIRT1-dependent deacetylation of PERK at lysine 1101, which suppresses downstream ATF4 transcription. Because ATF4 normally drives protective HSPA5 expression, nodosin-treated cells lose HSPA5 chaperone shielding. Consequently, the E3 ligase MARCHF1 binds GPX4 and catalyzes K48-linked polyubiquitination at lysine 191, driving proteasomal degradation and inducing selective ferroptosis.
Preclinical studies indicate that quiescent hepatocytes maintain robust alternative antioxidant pathways that protect against moderate lipid peroxidation. Furthermore, the SIRT1-PERK-HSPA5-MARCHF1 axis operates with heightened sensitivity in highly stressed, proliferative myofibroblasts. However, translational applications require precise dosing and targeted delivery systems to ensure therapeutic safety across diverse clinical liver pathologies.
Disclaimer: This content is for informational and educational purposes only... Refer to the latest local and national guidelines for clinical practice.
References
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


A preclinical breakthrough reveals how nodosin drives hepatic stellate cell ferroptosis via the SIRT1-PERK-HSPA5-MARCHF1 axis, degrading GPX4 and reversing experimental liver fibrosis. This discovery uncovers fresh therapeutic avenues for managing chronic hepatic fibrogenesis.
Today
A large UK cohort study of 1.4 million women shows that pregnancy complications, including gestational diabetes, hypertension, and preterm birth, substantially raise long-term risks for hypertension, chronic kidney disease, type 2 diabetes, and cardiovascular events, highlighting early postpartum intervention needs.
Today

A cross-sectional study reveals that hypertension knowledge, illness perceptions, depression, and substance use significantly predict self-care behaviors. Addressing mental health and cognitive beliefs in primary care is crucial for optimal blood pressure control and cardiovascular disease prevention.
Today
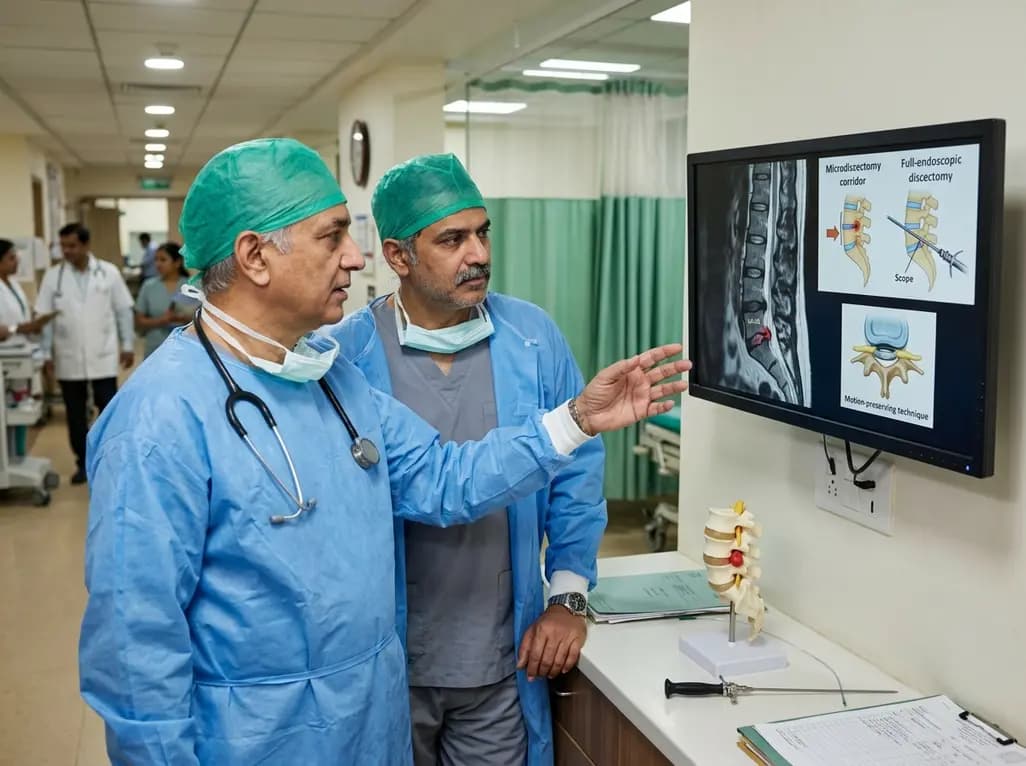
This clinical overview examines recent advancements in lumbar disc herniation surgery, comparing microscopic and full-endoscopic discectomy while highlighting biportal techniques and emerging motion-preserving innovations for spine specialists.
Today
Small interfering RNA (siRNA) therapeutics harness RNA interference to degrade disease-causing messenger RNA. By delivering hepatocyte-targeted, durable gene silencing, agents targeting PCSK9, LPA, and angiotensinogen offer potent cardiovascular risk reduction with infrequent, biannual dosing schedules.
Today