
Loading, please wait...
Loading, please wait...

Pancreatic islets maintain systemic metabolic equilibrium by orchestrating rapid hormone release in response to circulating fuels. Although science has long examined canonical ion channels, recent research provides compelling evidence that chloride cotransport fundamentally shapes islet physiology. Specifically, experimental investigations into beta-cell Kcc2, encoded by the Slc12a5 gene, demonstrate that this transporter regulates nutrient sensitivity and whole-body metabolic adaptation. Consequently, understanding these chloride pathways helps clinicians and researchers unravel novel dimensions of endocrine function and metabolic pathophysiology.
Pancreatic beta-cells depend on finely tuned bioelectrical networks to sense circulating fuel molecules and release appropriate amounts of insulin. Traditionally, potassium and calcium channels have dominated our conceptual framework of stimulus-secretion coupling. However, recent findings emphasize that intracellular chloride regulation is equally critical for fine-tuning the membrane potential. The solute carrier Slc12a5 encodes Kcc2, a potassium-chloride cotransporter originally characterized for its actions within mature neurons. In endocrine cells, Kcc2 extrudes chloride to prevent cellular hyperexcitability and preserve physiological stimulus-secretion dynamics.
When geneticists selectively eliminate beta-cell Kcc2 in murine models, the resting electrochemical balance within islet cells shifts markedly. Isolated islets lacking this transporter demonstrate elevated glucose-stimulated insulin secretion during early life. Because intracellular chloride accumulates without functional extrusion, membrane depolarization occurs more readily when nutrient levels rise. Therefore, the loss of this cotransporter accelerates secretory responses under specific metabolic conditions. Furthermore, this alteration highlights that chloride homeostasis serves as an essential regulatory checkpoint. Understanding these cellular mechanisms allows diabetologists to appreciate how subtle channel variations influence glycemic regulation across life stages.
The selective deletion of Kcc2 produces distinct systemic phenotypes between sexes, revealing complex neuroendocrine integration. Female knockout mice exhibit significantly lower body weight at weaning compared to wild-type littermates. However, as these female animals mature into adulthood, they develop increased lean mass without accumulating excess adiposity. In contrast, male knockout mice show reduced body-weight gain throughout post-weaning development, despite consuming larger amounts of food. This striking phenotypic divergence underscores the sexually dimorphic nature of metabolic compensation.
These divergent responses suggest that endocrine signals interact with sex steroids to direct peripheral nutrient utilization. In females, heightened insulin sensitivity or altered anabolic signaling may prioritize lean tissue accretion. Meanwhile, male animals appear to expend more energy or experience shifts in nutrient absorption efficiency. In addition, the dissociation between body weight and food intake indicates modified hypothalamic or autonomic communication. Thus, targeting beta-cell chloride transport triggers integrated systemic changes that transcend local islet output. Recognizing these sex differences provides meaningful insights into how baseline biological variation impacts metabolic disease vulnerability and tissue partitioning.
Beyond altering tissue composition, deleting Kcc2 from beta-cells reshapes feeding behavior in an age- and sex-dependent manner. Detailed feeding microstructure analyses demonstrate that male knockout mice alter their meal patterning substantially. Specifically, these animals adjust meal size, meal frequency, and the inter-meal intervals during early adulthood. Although they consume more total energy, they fail to gain commensurate body weight. Therefore, metabolic efficiency decreases as a compensatory adaptation to altered endocrine signaling.
These behavioral shifts suggest an active dialogue between pancreatic islets and central appetite-regulating centers. Peripheral hyperinsulinemia or altered postprandial hormone dynamics can directly alter vagal afferent firing or hypothalamic neuropeptide expression. Consequently, central nervous system circuits receive distorted feedback regarding peripheral energy status. Furthermore, meal microstructure reflects satiety cues and stomach emptying kinetics, which respond dynamically to circulating endocrine signals. Accordingly, these experimental findings indicate that islet-specific ion transport disruption reverberates throughout whole-body energy balance circuits. Studying these behavioral patterns helps researchers understand why metabolic disorders frequently present with concurrent abnormalities in appetite regulation and energy expenditure.
Morphometric evaluation reveals striking structural alterations in the endocrine pancreas of mice lacking Kcc2. Pancreatic sections show a significant increase in mean islet area across knockout animals. Notably, this islet enlargement occurs without disrupting the relative proportion of insulin-, glucagon-, or somatostatin-producing cells. This indicates that beta-cell hyperplasia or hypertrophy drives morphological expansion rather than endocrine lineage transdifferentiation. Thus, beta-cells adapt structurally to chronic alterations in ion transport and functional workload.
In parallel with these structural changes, glycemic regulation exhibits distinct age- and sex-dependent trajectories. Young male knockout mice display enhanced glucose tolerance during intraperitoneal challenges, mirroring their elevated in vitro insulin release. However, as animals transition into mature adulthood, these physiological advantages diminish or evolve into altered insulin sensitivity patterns. Additionally, fasting plasma insulin concentrations fluctuate widely based on animal age and fasting duration, while baseline fasting glycemia remains well defended. These dynamic adaptations underscore that beta-cell capacity is not static. Instead, islets continuously balance functional compensation against metabolic demand over time.
While glucose-stimulated insulin release increases in young knockout mice, responses to amino acid stimulation show the opposite pattern. Both male and female knockout mice exhibit significantly attenuated glycemic responses during alanine challenge tests. Under normal physiological circumstances, alanine transport into beta-cells induces electrogenic inward currents and fuels mitochondrial metabolism, triggering hormone secretion. However, the disruption of chloride homeostasis in Kcc2-deficient cells appears to compromise this specific nutrient-sensing pathway.
This selective impairment reveals that different nutrient secretagogues rely on distinct bioelectrical and biochemical cascades. Glucose primarily drives oxidative phosphorylation and closure of ATP-sensitive potassium channels. Conversely, amino acid signaling interfaces more directly with membrane transporters and electrogenic substrate entry. Therefore, altering intracellular chloride alters the membrane potential threshold differently for sugars versus amino acids. Furthermore, these divergent responses prove that enhanced glucose sensitivity does not confer universal secretagogue responsiveness. In clinical practice, recognizing that specific nutrients engage distinct secretory mechanisms assists clinicians in evaluating complex metabolic defects and designing tailored nutritional interventions.
Preclinical findings concerning Kcc2 illuminate important translational horizons for human diabetes management and endocrinology. Although Kcc2 was historically considered exclusive to neurons, its documented presence in human and rodent islets underscores unexplored cellular pharmacology. Ion channel modulators already represent a cornerstone of antidiabetic therapy, as demonstrated by sulfonylureas and glinides. Therefore, investigating islet chloride extruders provides fresh opportunities to develop highly targeted secretagogues with reduced hypoglycemia risk.
Moreover, the profound age- and sex-specific divergence observed in this knockout model mirrors the heterogeneity seen among diabetic patients. In clinical settings, individuals often present with widely differing insulin dynamics, body composition profiles, and feeding behaviors despite similar diagnostic criteria. Incorporating knowledge of non-canonical ion channels enriches our understanding of metabolic adaptation, beta-cell reserve, and endocrine plasticity. Consequently, continuing investigation into islet chloride regulation will undoubtedly refine our conceptual models of metabolic syndrome and type 2 diabetes pathogenesis.
Intracellular chloride balance directly modulates the resting membrane potential and electrical excitability of pancreatic endocrine tissue. When beta-cell Kcc2 is deleted, chloride extrusion decreases, which alters the depolarizing threshold required for voltage-gated calcium entry. Consequently, islets exhibit enhanced early insulin release in response to glucose stimulation. However, this electrophysiological shift attenuates compensatory secretory responses to amino acid challenges such as alanine, illustrating a complex, nutrient-specific modulation of endocrine stimulus-secretion coupling.
Sex hormones and divergent energetic demands substantially influence endocrine adaptations to altered chloride transport. Female knockout mice maintain stable systemic glucose handling while adjusting somatic growth and lean muscle distribution. Conversely, male mice display altered meal patterning, hyperphagia, and enhanced glucose clearance alongside age-dependent shifts in exogenous insulin sensitivity. These sex-specific divergences demonstrate that peripheral beta-cell signaling dynamically interfaces with systemic metabolic circuits governed by dimorphic neuroendocrine and steroid hormone pathways.
Pancreatic beta-cells rely on tightly regulated ion fluxes to coordinate postprandial insulin release and sustain normal glucose control. Uncovering the roles of non-canonical channels like Kcc2 reveals that chloride dynamics critically modulate beta-cell expansion and nutrient sensitivity. In clinical contexts, these molecular pathways could inspire novel antidiabetic therapies aimed at optimizing endogenous insulin secretion. Furthermore, identifying ion channel defects helps clinicians contextualize individual variations in insulin resistance, secretory dysfunction, and therapeutic responses.
Disclaimer: This content is for informational and educational purposes only... Refer to the latest local and national guidelines for clinical practice.
References
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.
Targeted deletion of the chloride cotransporter Kcc2 (Slc12a5) in pancreatic beta-cells triggers distinct sex- and age-dependent metabolic shifts, including altered feeding microstructure, enhanced insulin release in young mice, and modified glucose tolerance, highlighting chloride's role in islet biology.
Today
A novel high-permeation vesicle-in-gel platform enhances transdermal fulvestrant delivery, achieving sustained release, robust intradermal retention, and superior antitumor efficacy in breast cancer models while circumventing the drawbacks of painful intramuscular depot injections.
Today
A landmark ten-year WHO report reveals how socioeconomic status, geography, and gender disparities profoundly aggravate chronic disease risks, diagnosis delays, and therapeutic outcomes globally. Discover actionable primary care strategies to bridge screening divides and optimize vulnerable patient management.
Today
A recent single-institution analysis identifies advanced patient age and progressive systemic disease as primary negative prognostic factors for five-year survival in chondrosarcoma, underscoring the urgent necessity of precise risk stratification, clear surgical margins, and dedicated oncologic follow-up.
Today
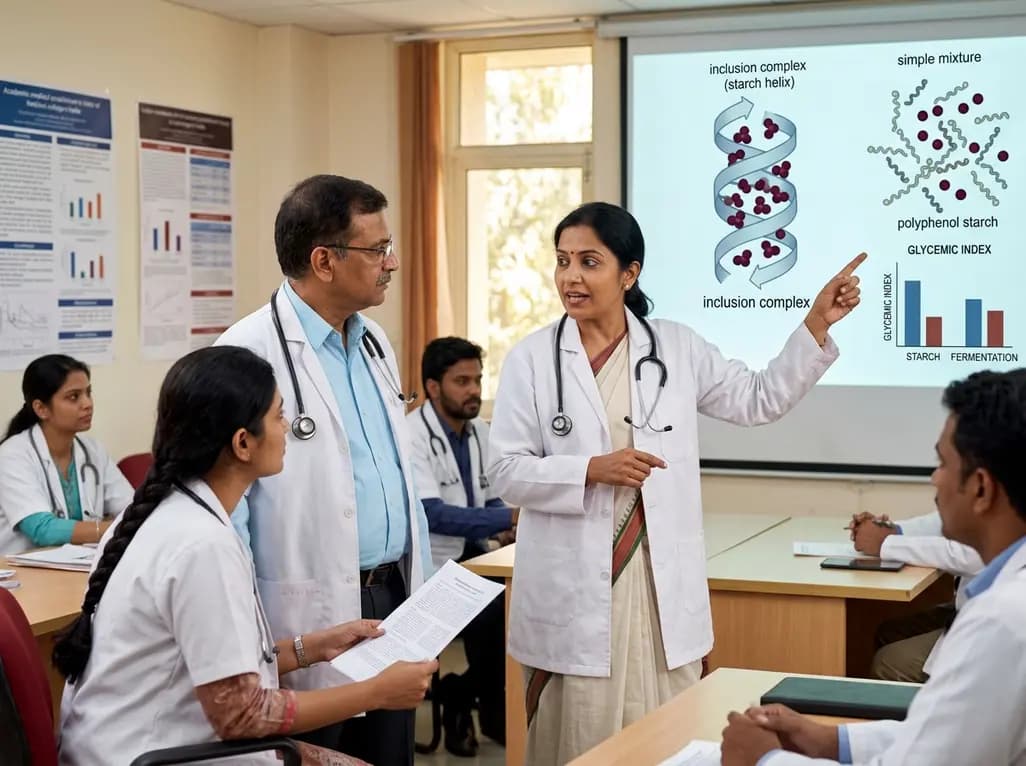
A critical review evaluates whether starch-polyphenol complexes meet structural and metabolic criteria to qualify as type 5 resistant starch. The proposed tiered evidence framework distinguishes true inclusion complexes from simple mixtures, clarifying mechanisms for glycemic modulation and gut fermentation.
Yesterday