Loading, please wait...
Loading, please wait...

Acute respiratory distress syndrome presents severe therapeutic challenges across intensive care units worldwide. Recent molecular studies identify acute lung injury ferroptosis as a pivotal driver of alveolar-capillary barrier degradation during septic inflammation. Lipopolysaccharide exposure triggers extensive cellular lipid peroxidation, compromising gas exchange and escalating mortality. Consequently, uncovering novel molecular pathways that protect alveolar epithelial cells from iron-mediated destruction is essential for advancing critical care medicine.
Alveolar epithelial cells serve as the primary defensive barrier preserving pulmonary gas exchange and fluid balance. During severe systemic sepsis, exposure to bacterial endotoxin precipitates uncontrolled production of reactive oxygen species within lung parenchymal cells. Furthermore, disruptions in cellular iron transport promote labile iron accumulation, which directly catalyzes lethal lipid peroxidation across cellular membranes. This iron-dependent non-apoptotic cell death pathway, termed ferroptosis, specifically degrades the structural integrity of fragile type I and type II pneumocytes. As epithelial integrity collapses, alveolar hyperpermeability permits protein-rich edema fluid to inundate distal airspaces, resulting in refractory hypoxemic respiratory failure. Additionally, dying alveolar cells release diverse danger-associated molecular patterns that amplify inflammatory leukocyte recruitment. Despite standard lung-protective ventilation, pharmacotherapies halting this rapid epithelial destruction remain scarce in clinical practice. Therefore, targeting the biochemical initiators of ferroptotic cell death represents a compelling therapeutic strategy. Understanding how endogenous cellular checkpoints govern lipid hydroperoxide clearance provides critical mechanistic pathways to rescue compromised respiratory tissues.
B-cell receptor-associated protein 31 operates as a key chaperone localized within the endoplasmic reticulum membrane. While scientists previously recognized BAP31 for regulating protein sorting and apoptotic trafficking, its influence on ferroptosis remained largely unexplored. To investigate this relationship, researchers constructed lipopolysaccharide-induced acute lung injury models in both wild-type mice and transgenic mice overexpressing BAP31. Subsequent transcriptomic profiling of lung tissues harvested twenty-four hours after endotoxin challenge demonstrated a pronounced reduction in baseline BAP31 expression. Notably, this pathological downregulation coincided directly with amplified markers of lipid peroxidation and extensive histological tissue damage. Conversely, transgenic overexpression of BAP31 significantly mitigated pulmonary edema, histopathological lung injury, and pro-inflammatory cytokine release. Unbiased RNA sequencing revealed that ferroptosis-related gene clusters formed the primary molecular network responsible for BAP31-mediated cytoprotection. These notable findings establish that BAP31 acts as an endogenous defender, whose loss during acute inflammatory stress renders alveolar epithelial cells vulnerable to catastrophic ferroptotic demise.
To delineate the precise intracellular signaling mechanisms, investigators examined the interaction between BAP31 and central survival kinases. Their experimental results demonstrated that BAP31 overexpression directly activates protein kinase B through targeted phosphorylation. Once activated, phosphorylated AKT suppresses glycogen synthase kinase 3β by phosphorylating its inhibitory serine residue. Under uncontrolled septic conditions lacking BAP31, constitutively active GSK3β promotes the cytoplasmic phosphorylation and rapid proteasomal degradation of crucial antioxidant factors. Conversely, BAP31-mediated activation of the AKT pathway efficiently turns off GSK3β enzymatic activity. In vitro knock-down experiments verified that blocking AKT phosphorylation completely abolishes the protective benefits of BAP31, leading to accelerated lipid peroxidation in alveolar cells. Furthermore, pharmacological inhibition of GSK3β successfully rescued cells lacking BAP31, demonstrating that GSK3β inactivation constitutes an obligatory downstream event. Thus, the AKT/GSK3β signaling cascade represents a vital intracellular conduit through which BAP31 stabilizes cellular homeostasis and prevents lipid peroxide accumulation during acute lung injury.
The primary downstream effector governed by the AKT/GSK3β axis is nuclear factor erythroid 2-related factor 2. Under suppressed GSK3β activity, Nrf2 avoids targeted cytoplasmic proteasomal degradation and rapidly translocates into the nucleus. Within the nuclear compartment, Nrf2 binds directly to antioxidant response elements in promoter regions of essential survival genes. Specifically, this transcriptional activation markedly upregulates glutathione peroxidase 4 and heme oxygenase 1 expression. Glutathione peroxidase 4 serves as the premier enzymatic barrier against ferroptosis by reducing toxic phospholipid hydroperoxides to benign lipid alcohols. Concurrently, heme oxygenase 1 degrades intracellular free heme, mitigating iron-catalyzed oxidative chain reactions. Crucially, experimental inhibition of Nrf2 completely nullified the anti-ferroptotic and antioxidant efficacy provided by BAP31 overexpression. These decisive findings prove that BAP31 suppresses ferroptosis primarily through the Nrf2-driven transcription of GPX4 and HMOX1. Consequently, this signaling pathway maintains critical redox equilibrium within distressed alveolar epithelial monolayers.
Preserving the alveolar epithelial barrier is fundamental to improving survival in patients suffering from acute respiratory distress syndrome. When bacterial lipopolysaccharide or sepsis destroys epithelial integrity, massive fluid exudation impairs pulmonary compliance and gas exchange. In experimental models, ferroptotic destruction of alveolar epithelial cells dramatically increases microvascular permeability and lung wet-to-dry ratios. However, sustaining BAP31 expression protects the alveolar barrier, markedly diminishing alveolar protein extravasation and systemic neutrophil infiltration. From a clinical perspective, identifying BAP31 and the AKT/GSK3β/Nrf2 axis highlights promising molecular targets for pharmacological intervention. Although clinical critical care currently depends upon supportive low tidal volume mechanical ventilation, targeted therapies preventing epithelial cell death could revolutionize practice. Inhaled or systemic agonists targeting the Nrf2 pathway could potentially reduce ventilator duration and prevent progressive fibroproliferation in severe lung injury. Therefore, translational validation of anti-ferroptotic therapies offers profound hope for reducing intensive care unit morbidity and mortality worldwide.
B-cell receptor-associated protein 31 acts as an essential molecular chaperone within the endoplasmic reticulum. In acute lung injury, inflammatory insults significantly downregulate BAP31 expression, accelerating iron-mediated lipid peroxidation. However, sustained BAP31 expression effectively activates downstream survival cascades, specifically promoting AKT phosphorylation. This enzymatic activation halts lethal lipid peroxide accumulation and shields delicate type I and type II alveolar epithelial cells from catastrophic membrane rupture during severe bacterial endotoxin challenge.
This biochemical signaling cascade serves as a vital regulator of endogenous cellular antioxidant defenses. When AKT phosphorylates and inactivates glycogen synthase kinase 3β, Nrf2 escapes proteasomal degradation and swiftly moves into the nucleus. Once translocated, Nrf2 drives the transcription of critical antiferroptotic enzymes, including GPX4 and HMOX1. In intensive care medicine, stimulating this intracellular cascade neutralizes overwhelming reactive oxygen species, thereby stabilizing alveolar-capillary barrier architecture under septic stress.
Current clinical management of acute respiratory distress syndrome relies almost entirely on supportive mechanical ventilation and fluid restriction. Unfortunately, disease-modifying pharmacotherapies directly preventing alveolar cell death remain unavailable. Targeting ferroptotic pathways addresses the core molecular mechanism driving alveolar barrier disintegration and fluid leakage. Therefore, developing pharmacological therapies that mimic BAP31 or activate Nrf2 could preserve pulmonary function, diminish mechanical ventilation dependency, and reduce intensive care unit mortality.
Disclaimer: This content is for informational and educational purposes only and should not be considered medical advice. Always consult a qualified healthcare provider for diagnosis and treatment decisions. Refer to the latest local and national guidelines for clinical practice.
References
Read summarized clinical updates, watch expert medical content, and earn CME certifications right from your smartphone.
Open your smartphone camera to scan and install.


A breakthrough study demonstrates that BAP31 protects against acute lung injury by inhibiting ferroptosis in alveolar epithelial cells. Through activating the AKT/GSK3β/Nrf2 pathway and upregulating GPX4 and HMOX1, BAP31 preserves the alveolar-capillary barrier, offering novel avenues for critical care management.
Today
A recent longitudinal study evaluated the link between prenatal loss of control eating and cardiovascular health using Life's Essential 8 frameworks. While direct associations across pregnancy were non-significant, the high prevalence of dysregulated eating highlights critical implications for obstetric practice.
Today
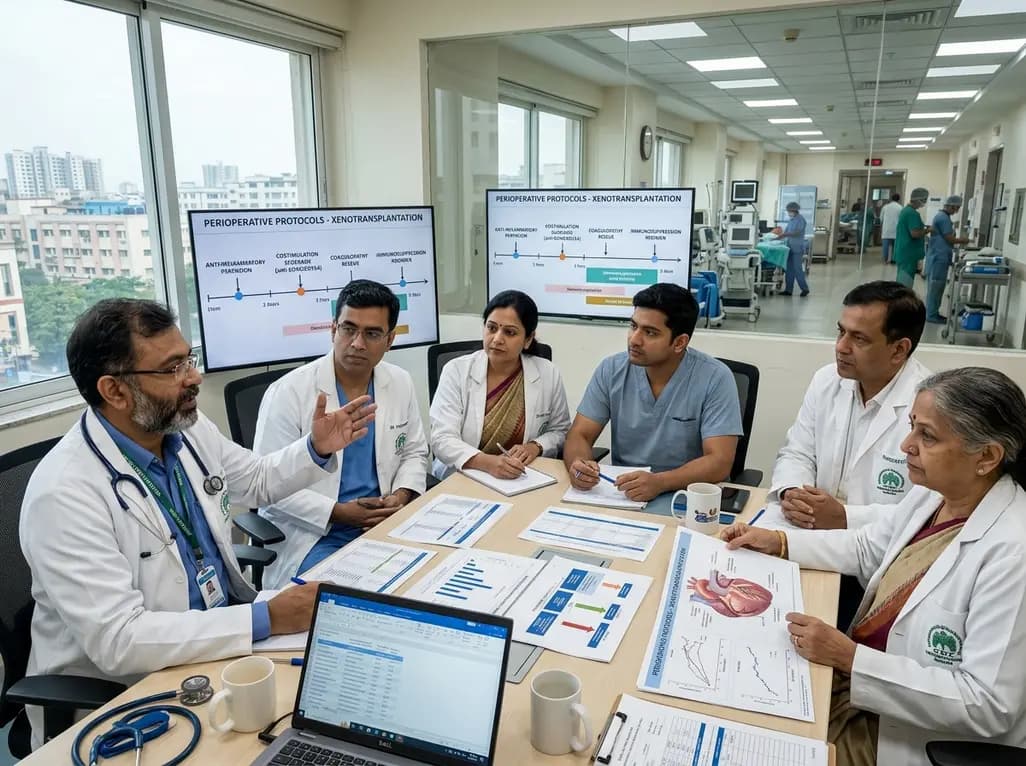
Explore modern perioperative management protocols for clinical xenotransplantation. Learn how costimulation blockade, anti-inflammatory perfusion, and coagulopathy rescue overcome interspecies hurdles.
Today
A retrospective study evaluating a locally deployed vision language model (Qwen3-VL) reveals that smartphone-captured radiographs and retrieval-augmented generation can paradoxically reduce diagnostic accuracy in bone tumor referrals due to optical noise and textual hallucinations.
Today
At Homa CME 2026 at Hyderabad's T-Hub, healthcare leaders emphasized moving beyond routine metrics through digital calculators and precision stratification. Discover why characterizing individual insulin secretion, resistance patterns, and vascular risks transforms chronic metabolic care in India.
Today